return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-270.170853
Energy at 298.15K-270.177771
HF Energy-269.183551
Nuclear repulsion energy262.062000
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3252 3080 14.53      
2 A1 3233 3062 1.42      
3 A1 3217 3047 8.15      
4 A1 3208 3038 6.13      
5 A1 1718 1627 1.38      
6 A1 1552 1470 12.62      
7 A1 1516 1436 3.34      
8 A1 1286 1218 5.04      
9 A1 1205 1141 1.80      
10 A1 1086 1029 2.51      
11 A1 992 940 0.06      
12 A1 863 817 0.87      
13 A1 547 518 0.65      
14 A2 1002 949 0.00      
15 A2 899 852 0.00      
16 A2 489 464 0.00      
17 A2 410 389 0.00      
18 B1 982 930 1.53      
19 B1 933 884 2.22      
20 B1 776 735 112.14      
21 B1 636 603 38.74      
22 B1 550 521 1.76      
23 B1 490 464 18.51      
24 B1 215 203 2.24      
25 B2 3319 3144 5.28      
26 B2 3239 3067 30.99      
27 B2 3220 3049 1.39      
28 B2 1699 1610 1.03      
29 B2 1545 1463 11.12      
30 B2 1382 1309 0.32      
31 B2 1286 1218 0.16      
32 B2 1172 1110 5.59      
33 B2 1157 1096 1.36      
34 B2 1004 951 1.60      
35 B2 648 614 0.00      
36 B2 364 344 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 25545.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24193.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18837 0.09163 0.06164

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.977
C2 0.000 1.204 0.246
C3 0.000 1.198 -1.121
C4 0.000 0.000 -1.820
C5 0.000 -1.198 -1.121
C6 0.000 -1.204 0.246
C7 0.000 0.000 2.387
H8 0.000 2.147 0.787
H9 0.000 2.139 -1.663
H10 0.000 0.000 -2.905
H11 0.000 -2.139 -1.663
H12 0.000 -2.147 0.787
H13 0.000 0.927 2.946
H14 0.000 -0.927 2.946

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.40872.41512.79672.41511.40871.41082.15503.39723.88163.39722.15502.17692.1769
C21.40871.36622.39122.76362.40852.45701.08672.12503.37303.84963.39432.71483.4402
C32.41511.36621.38732.39602.76363.70682.13031.08612.14933.38083.85034.07584.5884
C42.79672.39121.38731.38732.39124.20753.37712.14481.08492.14483.37714.85574.8557
C52.41512.76362.39601.38731.36623.70683.85033.38082.14931.08612.13034.58844.0758
C61.40872.40852.76362.39121.36622.45703.39433.84963.37302.12501.08673.44022.7148
C71.41082.45703.70684.20753.70682.45702.67754.58025.29244.58022.67751.08231.0823
H82.15501.08672.13033.37713.85033.39432.67752.44974.27074.93634.29322.48003.7562
H93.39722.12501.08612.14483.38083.84964.58022.44972.47364.27804.93634.76575.5356
H103.88163.37302.14931.08492.14933.37305.29244.27072.47362.47364.27075.92435.9243
H113.39723.84963.38082.14481.08612.12504.58024.93634.27802.47362.44975.53564.7657
H122.15503.39433.85033.37712.13031.08672.67754.29324.93634.27072.44973.75622.4800
H132.17692.71484.07584.85574.58843.44021.08232.48004.76575.92435.53563.75621.8537
H142.17693.44024.58844.85574.07582.71481.08233.75625.53565.92434.76572.48001.8537

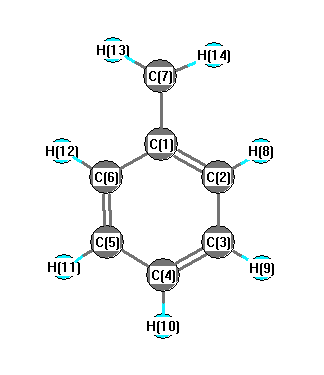
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.991 C1 C2 H8 118.873
C1 C6 C5 120.991 C1 C6 H12 118.873
C1 C7 H13 121.091 C1 C7 H14 121.091
C2 C1 C6 117.492 C2 C1 C7 121.254
C2 C3 C4 120.548 C2 C3 H9 119.687
C3 C2 H8 120.136 C3 C4 C5 119.432
C3 C4 H10 120.284 C4 C3 H9 119.765
C4 C5 C6 120.548 C4 C5 H11 119.765
C5 C1 C7 150.262 C5 C4 H10 120.284
C5 C6 H12 120.136 C6 C5 H11 119.687
H13 C7 H14 117.818
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability