return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-307.602792
Energy at 298.15K-307.608045
HF Energy-306.394545
Nuclear repulsion energy314.012902
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3265 3093 0.00      
2 Ag 3244 3072 0.00      
3 Ag 3229 3058 0.00      
4 Ag 1634 1548 0.00      
5 Ag 1548 1466 0.00      
6 Ag 1399 1325 0.00      
7 Ag 1329 1259 0.00      
8 Ag 1202 1138 0.00      
9 Ag 1112 1053 0.00      
10 Ag 983 931 0.00      
11 Ag 866 820 0.00      
12 Ag 716 678 0.00      
13 Ag 661 626 0.00      
14 Au 814 771 9.96      
15 Au 769 728 26.94      
16 Au 667 631 110.28      
17 Au 448 424 3.23      
18 Au 287 271 2.37      
19 Au 81 77 6.53      
20 Bg 854 809 0.00      
21 Bg 803 760 0.00      
22 Bg 699 662 0.00      
23 Bg 593 562 0.00      
24 Bg 510 483 0.00      
25 Bu 3265 3093 24.64      
26 Bu 3244 3073 11.48      
27 Bu 3230 3059 14.41      
28 Bu 1633 1547 32.02      
29 Bu 1523 1442 10.83      
30 Bu 1347 1276 19.38      
31 Bu 1276 1209 18.42      
32 Bu 1136 1076 39.88      
33 Bu 1063 1006 3.66      
34 Bu 969 918 5.39      
35 Bu 846 802 4.86      
36 Bu 443 420 3.70      

Unscaled Zero Point Vibrational Energy (zpe) 23844.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22582.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.17419 0.07203 0.05096

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.347 1.096 0.000
C2 2.169 -0.157 0.000
C3 1.347 -1.240 0.000
C4 -1.347 -1.096 0.000
C5 -2.169 0.157 0.000
C6 -1.347 1.240 0.000
C7 0.038 0.731 0.000
C8 -0.038 -0.731 0.000
H9 1.768 2.097 0.000
H10 3.254 -0.169 0.000
H11 1.646 -2.282 0.000
H12 -1.768 -2.097 0.000
H13 -3.254 0.169 0.000
H14 -1.646 2.282 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.49942.33643.47283.63912.69711.35852.29291.08622.28923.39134.46084.69333.2191
C21.49941.35993.63914.34993.78352.30912.28072.29031.08512.18794.38905.43334.5285
C32.33641.35992.69713.78353.66142.36631.47523.36412.18771.08393.23014.81194.6219
C43.47283.63912.69711.49942.33642.29291.35854.46084.69333.21911.08622.28923.3913
C53.63914.34993.78351.49941.35992.28072.30914.38905.43334.52852.29031.08512.1879
C62.69713.78353.66142.33641.35991.47522.36633.23014.81194.62193.36412.18771.0839
C71.35852.30912.36632.29292.28071.47521.46462.20413.33993.41553.35603.33992.2893
C82.29292.28071.47521.35852.30912.36631.46463.35603.33992.28932.20413.33993.4155
H91.08622.29033.36414.46084.38903.23012.20413.35602.71074.38105.48605.37943.4189
H102.28921.08512.18774.69335.43334.81193.33993.33992.71072.65495.37946.51735.4793
H113.39132.18791.08393.21914.52854.62193.41552.28934.38102.65493.41895.47935.6274
H124.46084.38903.23011.08622.29033.36413.35602.20415.48605.37943.41892.71074.3810
H134.69335.43334.81192.28921.08512.18773.33993.33995.37946.51735.47932.71072.6549
H143.21914.52854.62193.39132.18791.08392.28933.41553.41895.47935.62744.38102.6549

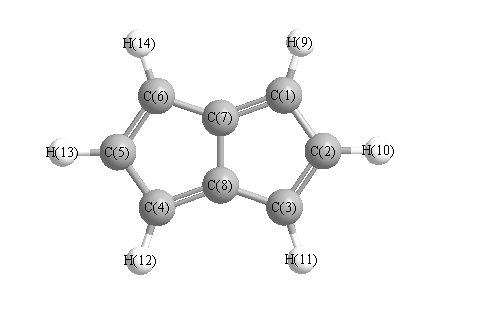
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.503 C1 C2 H10 123.900
C1 C7 C6 144.239 C1 C7 C8 108.558
C2 C1 C7 107.692 C2 C1 H9 123.920
C2 C3 C8 107.045 C2 C3 H11 126.728
C3 C2 H10 126.597 C3 C8 C4 144.239
C3 C8 C7 107.203 C4 C5 C6 109.503
C4 C5 H13 123.900 C4 C8 C7 108.558
C5 C4 C8 107.692 C5 C4 H12 123.920
C5 C6 C7 107.045 C5 C6 H14 126.728
C6 C5 H13 126.597 C6 C7 C8 107.203
C7 C1 H9 128.388 C7 C6 H14 126.227
C8 C3 H11 126.227 C8 C4 H12 128.388
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability