return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3C(OH)=NH (Ethaninidic acid)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-208.709095
Energy at 298.15K-208.715466
HF Energy-207.999002
Nuclear repulsion energy121.695394
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3776 3576 43.98      
2 A' 3581 3392 3.93      
3 A' 3208 3038 10.58      
4 A' 3099 2935 5.38      
5 A' 1760 1667 181.19      
6 A' 1524 1444 25.51      
7 A' 1481 1402 70.01      
8 A' 1419 1344 0.09      
9 A' 1303 1234 110.91      
10 A' 1134 1074 192.53      
11 A' 1039 984 34.46      
12 A' 892 845 1.06      
13 A' 558 529 45.17      
14 A' 430 408 2.40      
15 A" 3185 3017 5.13      
16 A" 1511 1431 11.64      
17 A" 1089 1031 7.19      
18 A" 865 819 24.74      
19 A" 644 610 169.56      
20 A" 525 497 13.95      
21 A" 115 109 1.22      

Unscaled Zero Point Vibrational Energy (zpe) 16570.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15693.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.36312 0.31021 0.17264

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.132 0.000
C2 0.939 -1.037 0.000
N3 0.249 1.382 0.000
O4 -1.287 -0.286 0.000
H5 1.978 -0.709 0.000
H6 0.755 -1.656 0.880
H7 0.755 -1.656 -0.880
H8 1.250 1.550 0.000
H9 -1.824 0.518 0.000

Atom - Atom Distances (Å)
  C1 C2 N3 O4 H5 H6 H7 H8 H9
C11.49981.27441.35292.14962.13082.13081.89021.8643
C21.49982.51552.34971.08931.09151.09152.60573.1710
N31.27442.51552.26702.71333.20263.20261.01482.2456
O41.35292.34972.26703.29242.61162.61163.13120.9668
H52.14961.08932.71333.29241.77971.77972.37363.9953
H62.13081.09153.20262.61161.77971.76043.36103.4858
H72.13081.09153.20262.61161.77971.76043.36103.4858
H81.89022.60571.01483.13122.37363.36103.36103.2422
H91.86433.17102.24560.96683.99533.48583.48583.2422

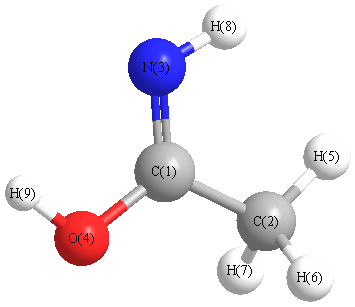
picture of Ethaninidic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 111.262 C1 C2 H6 109.623
C1 C2 H7 109.623 C1 N3 H8 110.812
C1 O4 H9 105.762 C2 C1 N3 129.942
C2 C1 O4 110.809 N3 C1 O4 119.249
H5 C2 H6 109.381 H5 C2 H7 109.381
H6 C2 H7 107.491
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability