return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-643.512870
Energy at 298.15K-643.520796
HF Energy-642.464886
Nuclear repulsion energy278.479316
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3581 3391 42.70      
2 A' 3229 3058 0.05      
3 A' 3115 2951 0.28      
4 A' 1684 1595 58.07      
5 A' 1496 1417 8.78      
6 A' 1416 1341 26.36      
7 A' 1179 1117 200.71      
8 A' 1032 977 27.52      
9 A' 917 868 133.15      
10 A' 763 722 9.32      
11 A' 700 663 275.44      
12 A' 513 485 49.58      
13 A' 483 458 13.44      
14 A' 306 290 4.58      
15 A" 3696 3501 50.44      
16 A" 3241 3069 0.14      
17 A" 1494 1415 2.40      
18 A" 1407 1333 266.96      
19 A" 1143 1082 1.14      
20 A" 1013 959 1.39      
21 A" 414 392 0.04      
22 A" 342 324 3.40      
23 A" 244 231 1.14      
24 A" 187 177 42.07      

Unscaled Zero Point Vibrational Energy (zpe) 16796.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15907.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.15732 0.14692 0.14391

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.661 -0.056 0.000
S2 0.110 -0.137 0.000
N3 0.528 1.481 0.000
O4 0.528 -0.695 1.272
O5 0.528 -0.695 -1.272
H6 -2.028 -1.082 0.000
H7 -1.990 0.465 0.896
H8 -1.990 0.465 -0.896
H9 1.039 1.722 0.838
H10 1.039 1.722 -0.838

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.77332.67462.61112.61111.08931.08761.08763.34043.3404
S21.77331.67071.45041.45042.33782.36112.36112.24092.2409
N32.67461.67072.52042.52043.61922.85862.85861.01131.0113
O42.61111.45042.52042.54362.88092.79713.51912.50853.2492
O52.61111.45042.52042.54362.88093.51912.79713.24922.5085
H61.08932.33783.61922.88092.88091.78821.78824.23954.2395
H71.08762.36112.85862.79713.51911.78821.79303.27993.7100
H81.08762.36112.85863.51912.79711.78821.79303.71003.2799
H93.34042.24091.01132.50853.24924.23953.27993.71001.6766
H103.34042.24091.01133.24922.50854.23953.71003.27991.6766

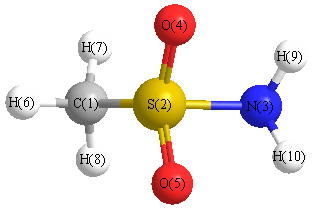
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 101.856 C1 S2 O4 107.767
C1 S2 O5 107.767 S2 C1 H6 107.069
S2 C1 H7 108.863 S2 C1 H8 108.863
S2 N3 H9 110.934 S2 N3 H10 110.934
N3 S2 O4 107.507 N3 S2 O5 107.507
O4 S2 O5 122.537 H6 C1 H7 110.455
H6 C1 H8 110.455 H7 C1 H8 111.026
H9 N3 H10 111.968
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability