return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11 (2-Methylbut-2-yl radical)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-196.507305
Energy at 298.15K-196.518313
HF Energy-195.740851
Nuclear repulsion energy182.432762
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3177 3009 26.84      
2 A 3169 3002 16.26      
3 A 3168 3000 48.87      
4 A 3161 2993 17.17      
5 A 3120 2955 22.46      
6 A 3111 2946 26.30      
7 A 3095 2931 20.49      
8 A 3082 2919 23.52      
9 A 3010 2851 48.30      
10 A 3006 2847 28.79      
11 A 2986 2828 39.55      
12 A 1553 1471 1.51      
13 A 1541 1460 10.00      
14 A 1533 1452 9.15      
15 A 1532 1451 23.49      
16 A 1522 1442 2.46      
17 A 1515 1435 4.48      
18 A 1508 1428 0.29      
19 A 1453 1376 7.57      
20 A 1446 1370 5.06      
21 A 1439 1363 11.25      
22 A 1410 1335 0.89      
23 A 1342 1271 0.81      
24 A 1310 1241 3.44      
25 A 1276 1209 4.41      
26 A 1144 1084 1.45      
27 A 1101 1043 0.17      
28 A 1071 1014 5.16      
29 A 1015 961 0.88      
30 A 1009 955 1.67      
31 A 967 916 0.81      
32 A 956 905 0.19      
33 A 781 740 1.14      
34 A 763 722 2.17      
35 A 463 438 0.65      
36 A 388 367 0.05      
37 A 315 298 2.33      
38 A 295 280 0.09      
39 A 225 214 0.85      
40 A 153 145 0.01      
41 A 146 138 0.06      
42 A 80 76 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 32667.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 30939.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.24963 0.11549 0.08532

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.566 1.477 0.065
H2 0.283 1.998 -0.382
H3 -1.481 1.902 -0.354
H4 -0.560 1.720 1.141
C5 2.050 -0.048 -0.150
H6 2.188 0.886 0.398
H7 2.904 -0.690 0.076
H8 2.071 0.179 -1.218
C9 0.741 -0.740 0.231
H10 0.713 -1.739 -0.220
H11 0.734 -0.912 1.323
C12 -0.506 -0.001 -0.154
C13 -1.792 -0.763 -0.089
H14 -1.683 -1.772 -0.494
H15 -2.139 -0.870 0.952
H16 -2.589 -0.256 -0.639

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15 H16
C11.09241.09281.10263.03552.83644.09153.20722.57843.47212.99591.49562.55753.48092.96072.7554
H21.09241.76691.76232.71342.33973.78272.68432.84283.76523.40252.16193.46614.25403.98393.6604
H31.09281.76691.76574.03883.88075.11244.04123.50134.25283.95422.14792.69583.68263.13412.4427
H41.10261.76231.76573.40602.96594.35243.85492.92763.92872.93842.15513.03244.01673.03973.3458
C53.03552.71344.03883.40601.09181.09261.09281.52842.15682.15542.55663.90814.12644.40874.6696
H62.83642.33973.88072.96591.09181.76121.76832.18293.07392.49052.89014.33544.78024.70265.0204
H74.09153.78275.11244.35241.09261.76121.76812.16942.44762.51293.48724.69984.74785.12225.5572
H83.20722.68434.04123.85491.09281.76831.76812.17102.55373.07172.79424.13344.29264.85144.7163
C92.57842.84283.50132.92761.52842.18292.16942.17101.09631.10521.49982.55322.73292.97203.4761
H103.47213.76524.25283.92872.15683.07392.44762.55371.09631.75002.12332.69132.41173.20323.6440
H112.99593.40253.95422.93842.15542.49052.51293.07171.10521.75002.13252.89723.14362.89703.9144
C121.49562.16192.14792.15512.55662.89013.48722.79421.49982.12332.13251.49542.15322.15492.1539
C132.55753.46612.69583.03243.90814.33544.69984.13342.55322.69132.89721.49541.09351.10241.0935
H143.48094.25403.68264.01674.12644.78024.74784.29262.73292.41173.14362.15321.09351.76451.7725
H152.96073.98393.13413.03974.40874.70265.12224.85142.97203.20322.89702.15491.10241.76451.7640
H162.75543.66042.44273.34584.66965.02045.55724.71633.47613.64403.91442.15391.09351.77251.7640

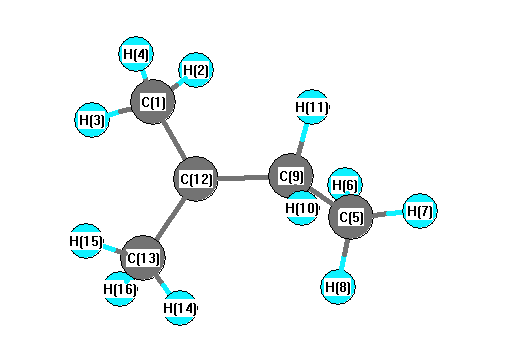
picture of 2-Methylbut-2-yl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 118.818 C1 C12 C13 117.532
H2 C1 H3 107.920 H2 C1 H4 106.805
H2 C1 C12 112.377 H3 C1 H4 107.078
H3 C1 C12 111.211 H4 C1 C12 111.185
C5 C9 H10 109.407 C5 C9 H11 108.793
C5 C9 C12 115.184 H6 C5 H7 107.473
H6 C5 H8 108.084 H6 C5 C9 111.755
H7 C5 H8 108.010 H7 C5 C9 110.627
H8 C5 C9 110.743 C9 C12 C13 116.955
H10 C9 H11 105.287 H10 C9 C12 108.754
H11 C9 C12 108.955 C12 C13 H14 111.611
C12 C13 H15 111.194 C12 C13 H16 111.661
H14 C13 H15 106.940 H14 C13 H16 108.295
H15 C13 H16 106.896
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability