return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Butene, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-195.959374
Energy at 298.15K-195.969025
HF Energy-195.179140
Nuclear repulsion energy174.305087
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3201 3032 33.57      
2 A 3186 3018 22.55      
3 A 3171 3003 26.47      
4 A 3163 2996 5.76      
5 A 3075 2912 20.48      
6 A 3069 2906 39.67      
7 A 3061 2899 26.25      
8 A 1749 1657 0.56      
9 A 1538 1457 16.88      
10 A 1536 1455 3.19      
11 A 1521 1441 3.23      
12 A 1458 1381 2.12      
13 A 1449 1372 5.01      
14 A 1447 1370 11.20      
15 A 1395 1321 1.31      
16 A 1261 1195 3.00      
17 A 1158 1096 6.69      
18 A 1087 1029 6.47      
19 A 990 938 2.27      
20 A 983 931 0.78      
21 A 785 743 1.54      
22 A 525 497 2.03      
23 A 388 367 0.31      
24 A 301 285 0.19      
25 A 3139 2973 18.98      
26 A 3133 2967 33.62      
27 A 3130 2964 12.96      
28 A 1531 1450 27.61      
29 A 1523 1442 0.85      
30 A 1508 1428 1.09      
31 A 1115 1056 0.00      
32 A 1069 1012 0.42      
33 A 1023 969 2.42      
34 A 808 765 16.82      
35 A 427 405 3.67      
36 A 261 247 1.88      
37 A 172 163 1.26      
38 A 104 99 0.47      
39 A 77 73 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 30257.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 28656.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.26854 0.11706 0.08538

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.504 0.499 0.000
H2 -1.969 -0.485 0.000
H3 -1.866 1.044 0.878
H4 -1.866 1.044 -0.878
C5 0.261 -2.096 0.000
H6 0.638 -2.629 -0.878
H7 0.638 -2.629 0.878
H8 -0.825 -2.182 0.000
C9 0.738 -0.674 0.000
H10 1.822 -0.554 0.000
C11 0.000 0.451 0.000
C12 0.663 1.803 0.000
H13 1.752 1.720 0.000
H14 0.364 2.384 0.879
H15 0.364 2.384 -0.879

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 C11 C12 H13 H14 H15
C11.08921.09531.09533.13883.89213.89212.76582.53073.48891.50462.52873.47662.79542.7954
H21.08921.76701.76702.75093.48793.48792.04612.71403.79202.18043.48784.32503.80143.8014
H31.09531.76701.75683.89354.78054.44603.50193.24154.11462.14622.78263.78352.60163.1396
H41.09531.76701.75683.89354.44604.78053.50193.24154.11462.14622.78263.78353.13962.6016
C53.13882.75093.89353.89351.09451.09451.08951.50022.19472.56063.92024.09704.56734.5673
H63.89213.48794.78054.44601.09451.75681.76452.14582.54573.26604.51874.57455.31985.0211
H73.89213.48794.44604.78051.09451.75681.76452.14582.54573.26604.51874.57455.02115.3198
H82.76582.04613.50193.50191.08951.76451.76452.17223.10802.75924.25394.67584.79974.7997
C92.53072.71403.24153.24151.50022.14582.14582.17221.09061.34562.47842.59953.20423.2042
H103.48893.79204.11464.11462.19472.54572.54573.10801.09062.08102.62692.27493.39613.3961
C111.50462.18042.14622.14622.56063.26603.26602.75921.34562.08101.50602.16282.15482.1548
C122.52873.48782.78262.78263.92024.51874.51874.25392.47842.62691.50601.09181.09551.0955
H133.47664.32503.78353.78354.09704.57454.57454.67582.59952.27492.16281.09181.77211.7721
H142.79543.80142.60163.13964.56735.31985.02114.79973.20423.39612.15481.09551.77211.7584
H152.79543.80143.13962.60164.56735.02115.31984.79973.20423.39612.15481.09551.77211.7584

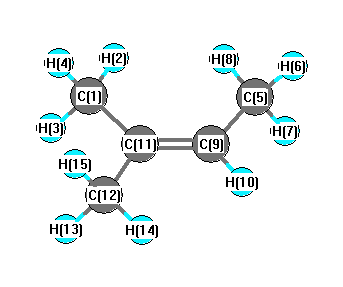
picture of 2-Butene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C11 C9 125.120 C1 C11 C12 114.269
H2 C1 H3 107.973 H2 C1 H4 107.973
H2 C1 C11 113.443 H3 C1 H4 106.629
H3 C1 C11 110.278 H4 C1 C11 110.278
C5 C9 H10 114.880 C5 C9 C11 128.168
H6 C5 H7 106.747 H6 C5 H8 107.782
H6 C5 C9 110.603 H7 C5 H8 107.782
H7 C5 C9 110.603 H8 C5 C9 113.064
C9 C11 C12 120.612 H10 C9 C11 116.952
C11 C12 H13 111.736 C11 C12 H14 110.866
C11 C12 H15 110.866 H13 C12 H14 108.221
H13 C12 H15 108.221 H14 C12 H15 106.746
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability