return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (3-Ethylthio-1-propene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-593.728909
Energy at 298.15K-593.739889
HF Energy-592.700865
Nuclear repulsion energy286.711063
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3289 3115 14.14      
2 A 3196 3027 4.01      
3 A 3178 3010 15.12      
4 A 3175 3007 20.08      
5 A 3087 2924 21.73      
6 A 3085 2921 26.78      
7 A 3063 2901 26.68      
8 A 1711 1621 16.19      
9 A 1543 1462 3.87      
10 A 1526 1445 2.46      
11 A 1504 1425 11.60      
12 A 1475 1397 6.87      
13 A 1452 1375 6.82      
14 A 1374 1301 7.53      
15 A 1344 1273 15.44      
16 A 1331 1260 14.91      
17 A 1113 1054 3.69      
18 A 1095 1037 2.06      
19 A 1029 975 3.80      
20 A 962 911 2.43      
21 A 787 745 2.49      
22 A 732 693 0.38      
23 A 515 487 0.20      
24 A 347 329 0.65      
25 A 253 239 0.21      
26 A 130 123 0.24      
27 A 3185 3016 22.22      
28 A 3141 2975 7.69      
29 A 3111 2946 14.27      
30 A 1533 1452 11.24      
31 A 1302 1233 0.23      
32 A 1214 1150 3.14      
33 A 1093 1035 0.88      
34 A 1013 959 21.73      
35 A 953 903 0.72      
36 A 900 852 48.13      
37 A 816 773 5.84      
38 A 544 515 13.84      
39 A 266 252 0.08      
40 A 112 106 0.51      
41 A 62 58 0.45      

Unscaled Zero Point Vibrational Energy (zpe) 30769.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29141.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.32666 0.04271 0.03859

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -3.797 -1.211 0.000
H2 -2.679 0.261 0.000
C3 -2.787 -0.818 0.000
H4 -1.915 -2.721 0.000
C5 -1.736 -1.646 0.000
H6 0.197 -1.699 0.882
H7 0.197 -1.699 -0.882
C8 -0.285 -1.262 0.000
S9 0.000 0.527 0.000
H10 2.159 -0.056 0.887
H11 2.159 -0.056 -0.887
C12 1.811 0.482 0.000
H13 3.451 1.886 0.000
H14 2.029 2.451 -0.884
H15 2.029 2.451 0.884
C16 2.359 1.903 0.000

Atom - Atom Distances (Å)
  H1 H2 C3 H4 C5 H6 H7 C8 S9 H10 H11 C12 H13 H14 H15 C16
H11.84851.08412.41282.10604.11944.11943.51184.17556.13106.13105.85767.88206.93776.93776.8986
H21.84851.08413.07802.12703.59033.59032.83662.69174.92804.92804.49486.34165.26675.26675.2984
C31.08411.08412.09311.33753.23383.23382.54023.09385.08155.08154.77756.79855.88695.88695.8206
H42.41283.07802.09311.08972.50662.50662.18733.76994.94804.94804.91317.07246.56396.56396.2966
C52.10602.12701.33751.08972.12582.12581.50082.78104.29944.29944.13636.27575.63425.63425.4190
H64.11943.59033.23382.50662.12581.76351.09632.40222.55893.11042.85284.92134.86834.53664.2927
H74.11943.59033.23382.50662.12581.76351.09632.40223.11042.55892.85284.92134.53664.86834.2927
C83.51182.83662.54022.18731.50081.09631.09631.81102.86602.86602.72674.88584.46374.46374.1243
S94.17552.69173.09383.76992.78102.40222.40221.81102.40522.40521.81133.70942.93302.93302.7313
H106.13104.92805.08154.94804.29942.55893.11042.86602.40521.77311.09372.49543.07222.51042.1595
H116.13104.92805.08154.94804.29943.11042.55892.86602.40521.77311.09372.49542.51043.07222.1595
C125.85764.49484.77754.91314.13632.85282.85282.72671.81131.09371.09372.15932.16972.16971.5232
H137.88206.34166.79857.07246.27574.92134.92134.88583.70942.49542.49542.15931.76781.76781.0924
H146.93775.26675.88696.56395.63424.86834.53664.46372.93303.07222.51042.16971.76781.76881.0916
H156.93775.26675.88696.56395.63424.53664.86834.46372.93302.51043.07222.16971.76781.76881.0916
C166.89865.29845.82066.29665.41904.29274.29274.12432.73132.15952.15951.52321.09241.09161.0916

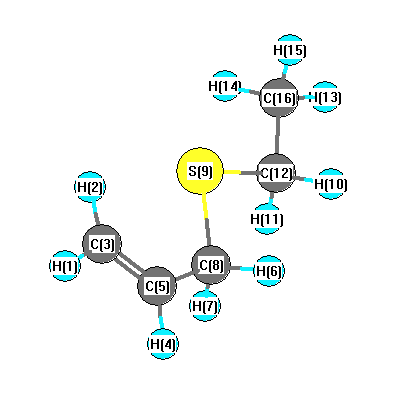
picture of 3-Ethylthio-1-propene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C3 H2 116.974 H1 C3 C5 120.485
H2 C3 C5 122.541 C3 C5 H4 118.815
C3 C5 C8 126.920 H4 C5 C8 114.265
C5 C8 H6 108.878 C5 C8 H7 108.878
C5 C8 S9 113.892 H6 C8 H7 107.089
H6 C8 S9 108.943 H7 C8 S9 108.943
C8 S9 C12 97.657 S9 C12 H10 109.277
S9 C12 H11 109.277 S9 C12 C16 109.684
H10 C12 H11 108.304 H10 C12 C16 110.136
H11 C12 C16 110.136 C12 C16 H13 110.192
C12 C16 H14 111.076 C12 C16 H15 111.076
H13 C16 H14 108.080 H13 C16 H15 108.080
H14 C16 H15 108.228
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability