Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -268.699087 |
| Energy at 298.15K | -268.705598 |
| HF Energy | -267.719653 |
| Nuclear repulsion energy | 222.424901 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3317 |
3142 |
1.18 |
|
|
|
| 2 |
A' |
3293 |
3119 |
0.60 |
|
|
|
| 3 |
A' |
3281 |
3108 |
3.45 |
|
|
|
| 4 |
A' |
3190 |
3021 |
9.89 |
|
|
|
| 5 |
A' |
3082 |
2919 |
22.52 |
|
|
|
| 6 |
A' |
1643 |
1556 |
9.28 |
|
|
|
| 7 |
A' |
1547 |
1465 |
7.40 |
|
|
|
| 8 |
A' |
1527 |
1446 |
18.49 |
|
|
|
| 9 |
A' |
1451 |
1374 |
6.28 |
|
|
|
| 10 |
A' |
1432 |
1356 |
1.10 |
|
|
|
| 11 |
A' |
1298 |
1230 |
26.11 |
|
|
|
| 12 |
A' |
1268 |
1200 |
2.92 |
|
|
|
| 13 |
A' |
1206 |
1143 |
25.88 |
|
|
|
| 14 |
A' |
1127 |
1068 |
1.81 |
|
|
|
| 15 |
A' |
1056 |
1000 |
22.13 |
|
|
|
| 16 |
A' |
1004 |
951 |
3.44 |
|
|
|
| 17 |
A' |
951 |
901 |
15.33 |
|
|
|
| 18 |
A' |
900 |
853 |
2.88 |
|
|
|
| 19 |
A' |
665 |
630 |
1.14 |
|
|
|
| 20 |
A' |
335 |
317 |
1.89 |
|
|
|
| 21 |
A" |
3158 |
2991 |
12.97 |
|
|
|
| 22 |
A" |
1511 |
1431 |
10.35 |
|
|
|
| 23 |
A" |
1070 |
1013 |
0.82 |
|
|
|
| 24 |
A" |
773 |
732 |
15.46 |
|
|
|
| 25 |
A" |
759 |
719 |
26.19 |
|
|
|
| 26 |
A" |
705 |
667 |
44.80 |
|
|
|
| 27 |
A" |
625 |
592 |
11.60 |
|
|
|
| 28 |
A" |
569 |
539 |
0.15 |
|
|
|
| 29 |
A" |
232 |
220 |
5.68 |
|
|
|
| 30 |
A" |
114 |
108 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21544.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20404.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.038 |
-0.238 |
0.000 |
| C2 |
0.000 |
0.645 |
0.000 |
| C3 |
0.505 |
-1.489 |
0.000 |
| C4 |
-1.189 |
-0.037 |
0.000 |
| C5 |
-0.861 |
-1.428 |
0.000 |
| C6 |
0.355 |
2.088 |
0.000 |
| H7 |
1.216 |
-2.301 |
0.000 |
| H8 |
-2.174 |
0.407 |
0.000 |
| H9 |
-1.540 |
-2.268 |
0.000 |
| H10 |
-0.554 |
2.691 |
0.000 |
| H11 |
0.943 |
2.350 |
0.884 |
| H12 |
0.943 |
2.350 |
-0.884 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3634 | 1.3602 | 2.2363 | 2.2412 | 2.4247 | 2.0700 | 3.2770 | 3.2818 | 3.3344 | 2.7362 | 2.7362 |
C2 | 1.3634 | | 2.1935 | 1.3705 | 2.2451 | 1.4859 | 3.1871 | 2.1874 | 3.2958 | 2.1198 | 2.1389 | 2.1389 | C3 | 1.3602 | 2.1935 | | 2.2312 | 1.3669 | 3.5807 | 1.0791 | 3.2825 | 2.1881 | 4.3127 | 3.9637 | 3.9637 | C4 | 2.2363 | 1.3705 | 2.2312 | | 1.4296 | 2.6264 | 3.3030 | 1.0809 | 2.2593 | 2.8010 | 3.3196 | 3.3196 | C5 | 2.2412 | 2.2451 | 1.3669 | 1.4296 | | 3.7206 | 2.2529 | 2.2570 | 1.0805 | 4.1310 | 4.2786 | 4.2786 | C6 | 2.4247 | 1.4859 | 3.5807 | 2.6264 | 3.7206 | | 4.4725 | 3.0369 | 4.7510 | 1.0908 | 1.0932 | 1.0932 | H7 | 2.0700 | 3.1871 | 1.0791 | 3.3030 | 2.2529 | 4.4725 | | 4.3393 | 2.7565 | 5.2965 | 4.7413 | 4.7413 | H8 | 3.2770 | 2.1874 | 3.2825 | 1.0809 | 2.2570 | 3.0369 | 4.3393 | | 2.7500 | 2.8006 | 3.7778 | 3.7778 | H9 | 3.2818 | 3.2958 | 2.1881 | 2.2593 | 1.0805 | 4.7510 | 2.7565 | 2.7500 | | 5.0570 | 5.3173 | 5.3173 | H10 | 3.3344 | 2.1198 | 4.3127 | 2.8010 | 4.1310 | 1.0908 | 5.2965 | 2.8006 | 5.0570 | | 1.7715 | 1.7715 | H11 | 2.7362 | 2.1389 | 3.9637 | 3.3196 | 4.2786 | 1.0932 | 4.7413 | 3.7778 | 5.3173 | 1.7715 | | 1.7672 | H12 | 2.7362 | 2.1389 | 3.9637 | 3.3196 | 4.2786 | 1.0932 | 4.7413 | 3.7778 | 5.3173 | 1.7715 | 1.7672 | |
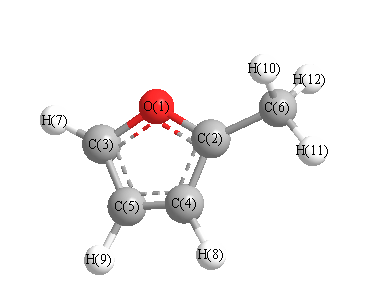 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.767 |
|
O1 |
C2 |
C6 |
116.573 |
| O1 |
C3 |
C5 |
110.537 |
|
O1 |
C3 |
H7 |
115.635 |
| C2 |
O1 |
C3 |
107.290 |
|
C2 |
C4 |
C5 |
106.578 |
| C2 |
C4 |
H8 |
125.913 |
|
C2 |
C6 |
H10 |
109.756 |
| C2 |
C6 |
H11 |
111.148 |
|
C2 |
C6 |
H12 |
111.148 |
| C3 |
C5 |
C6 |
73.493 |
|
C3 |
C5 |
H9 |
126.379 |
| C4 |
C2 |
C6 |
133.659 |
|
C4 |
C5 |
H9 |
127.794 |
| C5 |
C3 |
H7 |
133.827 |
|
C5 |
C4 |
H8 |
127.509 |
| H10 |
C6 |
H11 |
108.417 |
|
H10 |
C6 |
H12 |
108.417 |
| H11 |
C6 |
H12 |
107.863 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
