return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9NO (Butanamide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-287.132320
Energy at 298.15K-287.142940
HF Energy-286.106987
Nuclear repulsion energy238.992629
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3775 3575 29.23      
2 A 3638 3446 35.23      
3 A 3172 3005 43.61      
4 A 3167 2999 31.91      
5 A 3150 2983 3.88      
6 A 3119 2954 12.62      
7 A 3104 2940 19.07      
8 A 3075 2912 26.69      
9 A 3065 2903 13.08      
10 A 1812 1716 255.63      
11 A 1689 1600 97.49      
12 A 1547 1465 9.56      
13 A 1542 1461 10.23      
14 A 1531 1450 2.98      
15 A 1505 1426 9.58      
16 A 1452 1375 21.61      
17 A 1449 1372 60.48      
18 A 1385 1312 47.26      
19 A 1357 1285 25.39      
20 A 1288 1220 35.61      
21 A 1285 1217 1.76      
22 A 1168 1106 2.89      
23 A 1144 1084 1.28      
24 A 1112 1053 1.55      
25 A 1086 1029 3.41      
26 A 955 905 3.77      
27 A 908 860 0.93      
28 A 872 826 3.05      
29 A 761 720 8.68      
30 A 659 624 9.20      
31 A 630 596 35.33      
32 A 512 485 27.39      
33 A 438 415 8.54      
34 A 391 370 229.84      
35 A 351 332 4.92      
36 A 253 239 0.06      
37 A 187 177 8.70      
38 A 98 93 0.24      
39 A 35 33 3.51      

Unscaled Zero Point Vibrational Energy (zpe) 29330.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 27779.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.28879 0.06152 0.05268

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.720 -0.220 0.022
H2 -2.863 -0.504 1.068
H3 -3.520 0.474 -0.246
H4 -2.843 -1.120 -0.588
C5 -1.348 0.410 -0.192
H6 -1.234 0.722 -1.234
H7 -1.241 1.316 0.409
C8 -0.216 -0.545 0.164
H9 -0.260 -1.443 -0.463
H10 -0.329 -0.884 1.203
N11 2.188 -0.773 -0.157
H12 3.116 -0.398 -0.047
H13 2.061 -1.750 0.043
C14 1.153 0.106 0.041
O15 1.323 1.312 0.092

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 N11 H12 H13 C14 O15
C11.09361.09251.09361.52492.16192.16662.52892.78922.74854.94235.83945.02033.88624.3237
H21.09361.76501.76652.17233.07562.52492.79733.16202.56625.20446.08345.18204.18954.6661
H31.09251.76501.76502.17392.50302.51583.48183.78783.75875.84346.69636.01524.69574.9261
H41.09361.76651.76502.17562.52983.08062.79232.60603.09605.06156.02724.98504.22644.8713
C51.52492.17232.17392.17561.09441.09201.52312.16552.15823.72874.53894.04262.52942.8328
H62.16193.07562.50302.52981.09441.74762.14442.49623.05613.88694.64634.31302.77512.9399
H72.16662.52492.51583.08061.09201.74762.13863.05492.50994.05514.70474.52102.70732.5835
C82.52892.79733.48182.79231.52312.14442.13861.09591.09832.43623.34232.57941.52052.4131
H92.78923.16203.78782.60602.16552.49623.05491.09591.75842.55683.55892.39602.15603.2256
H102.74852.56623.75873.09602.15823.05612.50991.09831.75842.86273.69692.79452.12692.9638
N114.94235.20445.84345.06153.72873.88694.05512.43622.55682.86271.00701.00491.37262.2716
H125.83946.08346.69636.02724.53894.64634.70473.34233.55893.69691.00701.71692.02942.4825
H135.02035.18206.01524.98504.04264.31304.52102.57942.39602.79451.00491.71692.06643.1505
C143.88624.18954.69574.22642.52942.77512.70731.52052.15602.12691.37262.02942.06641.2195
O154.32374.66614.92614.87132.83282.93992.58352.41313.22562.96382.27162.48253.15051.2195

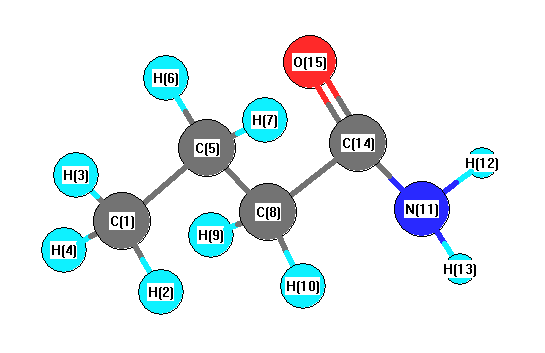
picture of Butanamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.164 C1 C5 H7 110.685
C1 C5 C8 112.133 H2 C1 H3 107.676
H2 C1 H4 107.731 H2 C1 C5 111.040
H3 C1 H4 107.683 H3 C1 C5 111.234
H4 C1 C5 111.302 C5 C8 H9 110.490
C5 C8 H10 109.767 C5 C8 C14 112.412
H6 C5 H7 106.133 H6 C5 C8 108.921
H7 C5 C8 108.604 C8 C14 N11 114.619
C8 C14 O15 123.072 H9 C8 H10 106.523
H9 C8 C14 109.917 H10 C8 C14 107.517
N11 C14 O15 122.300 H12 N11 H13 117.163
H12 N11 C14 116.194 H13 N11 C14 119.921
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability