Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -93.724185 |
| Energy at 298.15K | -93.725433 |
| HF Energy | -93.411450 |
| Nuclear repulsion energy | 28.580272 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3473 |
3289 |
12.70 |
|
|
|
| 2 |
A' |
3027 |
2867 |
337.98 |
|
|
|
| 3 |
A' |
2456 |
2326 |
379.91 |
|
|
|
| 4 |
A' |
1014 |
960 |
94.59 |
|
|
|
| 5 |
A' |
725 |
686 |
370.99 |
|
|
|
| 6 |
A" |
923 |
874 |
11.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5808.6 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 5501.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.107 |
0.629 |
0.000 |
| N2 |
0.107 |
-0.571 |
0.000 |
| H3 |
-0.676 |
1.405 |
0.000 |
| H4 |
-0.714 |
-1.177 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2002 | 1.1027 | 1.9830 |
N2 | 1.2002 | | 2.1260 | 1.0195 | H3 | 1.1027 | 2.1260 | | 2.5819 | H4 | 1.9830 | 1.0195 | 2.5819 | |
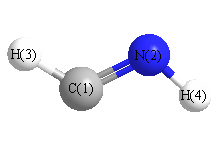 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
126.411 |
|
H3 |
C1 |
N2 |
134.750 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -93.733932 |
| Energy at 298.15K | -93.735253 |
| HF Energy | -93.418269 |
| Nuclear repulsion energy | 28.490471 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3588 |
3398 |
54.17 |
|
|
|
| 2 |
A' |
3073 |
2911 |
219.33 |
|
|
|
| 3 |
A' |
2482 |
2351 |
448.50 |
|
|
|
| 4 |
A' |
1223 |
1158 |
11.47 |
|
|
|
| 5 |
A' |
829 |
785 |
499.01 |
|
|
|
| 6 |
A" |
968 |
916 |
216.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6081.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 5759.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.005 |
0.643 |
0.000 |
| N2 |
-0.005 |
-0.569 |
0.000 |
| H3 |
0.913 |
1.251 |
0.000 |
| H4 |
-0.853 |
-1.125 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2120 | 1.1002 | 1.9612 |
N2 | 1.2120 | | 2.0377 | 1.0144 | H3 | 1.1002 | 2.0377 | | 2.9600 | H4 | 1.9612 | 1.0144 | 2.9600 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
123.253 |
|
H3 |
C1 |
N2 |
123.517 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
