return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,5-Hexadiene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.944713
Energy at 298.15K-233.954468
HF Energy-233.024902
Nuclear repulsion energy212.634789
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3277 3104 6.86      
2 A 3183 3015 0.19      
3 A 3171 3003 37.04      
4 A 3138 2972 21.05      
5 A 3064 2902 3.77      
6 A 1706 1616 4.86      
7 A 1520 1439 3.50      
8 A 1474 1396 4.12      
9 A 1385 1311 0.25      
10 A 1336 1265 1.09      
11 A 1269 1201 0.20      
12 A 1107 1049 0.02      
13 A 1069 1012 2.26      
14 A 1019 965 10.37      
15 A 970 919 0.12      
16 A 905 857 16.00      
17 A 790 748 0.50      
18 A 636 602 7.39      
19 A 428 406 0.00      
20 A 353 335 0.10      
21 A 95 90 0.01      
22 A 76 72 0.01      
23 B 3277 3104 24.78      
24 B 3183 3014 9.77      
25 B 3170 3003 9.30      
26 B 3119 2954 1.40      
27 B 3071 2909 41.91      
28 B 1709 1619 9.74      
29 B 1533 1451 11.60      
30 B 1475 1397 0.34      
31 B 1349 1277 0.74      
32 B 1330 1259 0.23      
33 B 1283 1215 0.27      
34 B 1202 1138 2.57      
35 B 1021 967 37.58      
36 B 979 928 3.84      
37 B 955 904 0.37      
38 B 905 857 73.25      
39 B 662 627 24.26      
40 B 440 416 1.17      
41 B 226 214 0.25      
42 B 111 105 0.15      

Unscaled Zero Point Vibrational Energy (zpe) 31482.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29817.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.40513 0.04756 0.04594

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.425 0.641 -0.321
C2 0.425 -0.641 -0.321
C3 0.425 1.875 -0.352
C4 -0.425 -1.875 -0.352
C5 0.486 2.779 0.633
C6 -0.486 -2.779 0.633
H7 -1.093 0.627 -1.192
H8 -1.062 0.648 0.570
H9 1.093 -0.627 -1.192
H10 1.062 -0.648 0.570
H11 1.046 2.014 -1.238
H12 -1.046 -2.014 -1.238
H13 1.133 3.647 0.568
H14 -0.115 2.676 1.532
H15 -1.133 -3.647 0.568
H16 0.115 -2.676 1.532

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.53801.49892.51552.51243.55071.09751.09512.16112.16032.21212.87683.50092.76934.43593.8371
C21.53802.51551.49893.55072.51242.16112.16031.09751.09512.87682.21214.43593.83713.50092.7693
C31.49892.51553.84441.33894.84322.13692.13712.72172.76051.09064.25122.11912.11755.81074.9347
C42.51551.49893.84444.84321.33892.72172.76052.13692.13714.25121.09065.81074.93472.11912.1175
C52.51243.55071.33894.84325.64223.23332.63463.91123.47572.09745.36871.08451.08626.62715.5406
C63.55072.51244.84321.33895.64223.91123.47573.23332.63465.36872.09746.62715.54061.08451.0862
H71.09752.16112.13692.72173.23333.91121.76202.52003.06172.55022.64204.14413.54544.62254.4480
H81.09512.16032.13712.76052.63463.47571.76203.06172.48873.09523.21823.71642.43574.29593.6552
H92.16111.09752.72172.13693.91123.23332.52003.06171.76202.64202.55024.62254.44804.14413.5454
H102.16031.09512.76052.13713.47572.63463.06172.48871.76203.21823.09524.29593.65523.71642.4357
H112.21212.87681.09064.25122.09745.36872.55023.09522.64203.21824.53962.43653.07536.32955.5257
H122.87682.21214.25121.09065.36872.09742.64203.21822.55023.09524.53966.32955.52572.43653.0753
H133.50094.43592.11915.81071.08456.62714.14413.71644.62254.29592.43656.32951.85187.63806.4763
H142.76933.83712.11754.93471.08625.54063.54542.43574.44803.65523.07535.52571.85186.47635.3566
H154.43593.50095.81072.11916.62711.08454.62254.29594.14413.71646.32952.43657.63806.47631.8518
H163.83712.76934.93472.11755.54061.08624.44803.65523.54542.43575.52573.07536.47635.35661.8518

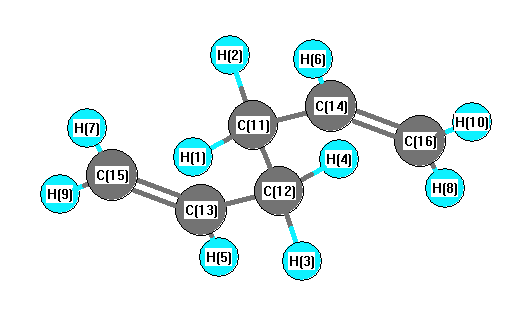
picture of 1,5-Hexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 111.846 C1 C2 H9 109.023
C1 C2 H10 109.107 C1 C3 C5 124.491
C1 C3 H11 116.465 C2 C1 C3 111.846
C2 C1 H7 109.023 C2 C1 H8 109.107
C2 C4 C6 124.491 C2 C4 H12 116.465
C3 C1 H7 109.807 C3 C1 H8 109.974
C3 C5 H13 121.597 C3 C5 H14 121.305
C4 C2 H9 109.807 C4 C2 H10 109.974
C4 C6 H15 121.597 C4 C6 H16 121.305
C5 C3 H11 119.029 C6 C4 H12 119.029
H7 C1 H8 106.954 H9 C2 H10 106.954
H13 C5 H14 117.098 H15 C6 H16 117.098
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability