return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((Z)-hexa-2,3,4-triene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-232.723187
Energy at 298.15K-232.729355
HF Energy-231.810602
Nuclear repulsion energy190.131744
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3188 3020 19.00      
2 A1 3164 2997 6.98      
3 A1 3070 2908 12.86      
4 A1 2174 2059 1.46      
5 A1 1533 1452 8.87      
6 A1 1442 1365 10.15      
7 A1 1401 1326 0.36      
8 A1 1146 1086 1.57      
9 A1 1086 1029 0.61      
10 A1 769 728 0.34      
11 A1 473 448 6.70      
12 A1 96 91 1.59      
13 A2 3142 2976 0.00      
14 A2 1518 1438 0.00      
15 A2 1067 1011 0.00      
16 A2 839 794 0.00      
17 A2 464 440 0.00      
18 A2 230 218 0.00      
19 A2 131 124 0.00      
20 B1 3142 2976 31.82      
21 B1 1519 1439 17.69      
22 B1 1064 1008 0.74      
23 B1 691 654 48.84      
24 B1 223 211 3.54      
25 B1 122 116 1.92      
26 B2 3188 3020 11.99      
27 B2 3165 2998 0.16      
28 B2 3070 2908 43.86      
29 B2 1705 1615 4.86      
30 B2 1522 1442 17.82      
31 B2 1440 1363 3.06      
32 B2 1298 1229 13.53      
33 B2 1121 1062 1.96      
34 B2 959 908 28.47      
35 B2 550 521 2.81      
36 B2 210 199 0.40      

Unscaled Zero Point Vibrational Energy (zpe) 25960.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24586.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.41604 0.04632 0.04234

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.000 2.179 1.631
H2 0.881 3.452 0.765
H3 -0.881 3.452 0.765
H4 0.000 -2.179 1.631
H5 0.881 -3.452 0.765
H6 -0.881 -3.452 0.765
C7 0.000 2.803 0.737
C8 0.000 -2.803 0.737
H9 0.000 2.486 -1.466
H10 0.000 -2.486 -1.466
C11 0.000 -0.636 -0.510
C12 0.000 0.636 -0.510
C13 0.000 1.962 -0.509
C14 0.000 -1.962 -0.509

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 H9 H10 C11 C12 C13 C14
H11.77421.77424.35725.76475.76471.09055.06143.11165.59843.53632.63842.15044.6611
H21.77421.76175.76476.90457.12561.09456.31722.58576.40384.37223.21412.14885.6318
H31.77421.76175.76477.12566.90451.09456.31722.58576.40384.37223.21412.14885.6318
H44.35725.76475.76471.77421.77425.06141.09055.59843.11162.63843.53634.66112.1504
H55.76476.90457.12561.77421.76176.31721.09456.40382.58573.21414.37225.63182.1488
H65.76477.12566.90451.77421.76176.31721.09456.40382.58573.21414.37225.63182.1488
C71.09051.09451.09455.06146.31726.31725.60642.22535.72913.65842.50001.50294.9258
C85.06146.31726.31721.09051.09451.09455.60645.72912.22532.50003.65844.92581.5029
H93.11162.58572.58575.59846.40386.40382.22535.72914.97113.26482.08171.09054.5498
H105.59846.40386.40383.11162.58572.58575.72912.22534.97112.08173.26484.54981.0905
C113.53634.37224.37222.63843.21413.21413.65842.50003.26482.08171.27242.59871.3263
C122.63843.21413.21413.53634.37224.37222.50003.65842.08173.26481.27241.32632.5987
C132.15042.14882.14884.66115.63185.63181.50294.92581.09054.54982.59871.32633.9250
C144.66115.63185.63182.15042.14882.14884.92581.50294.54981.09051.32632.59873.9250

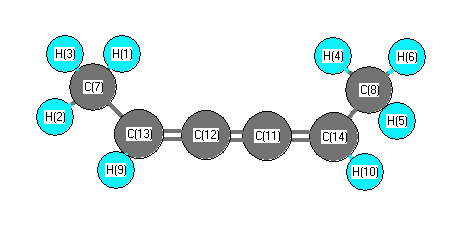
picture of (Z)-hexa-2,3,4-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C7 H2 108.582 H1 C7 H3 108.582
H1 C7 C13 111.039 H2 C7 H3 107.182
H2 C7 C13 110.669 H3 C7 C13 110.669
H4 C8 H5 108.582 H4 C8 H6 108.582
H4 C8 C14 111.039 H5 C8 H6 107.182
H5 C8 C14 110.669 H6 C8 C14 110.669
C7 C13 H9 117.320 C7 C13 C12 124.060
C8 C14 H10 117.320 C8 C14 C11 124.060
H9 C13 C12 118.620 H10 C14 C11 118.620
C11 C12 C13 179.956 C12 C11 C14 179.956
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability