return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,2-Butadiene, 3-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-194.737908
Energy at 298.15K-194.744955
HF Energy-193.972339
Nuclear repulsion energy157.073320
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3186 3017 18.44      
2 A1 3164 2996 5.52      
3 A1 3067 2905 43.82      
4 A1 2055 1947 11.82      
5 A1 1546 1464 0.18      
6 A1 1496 1417 4.33      
7 A1 1448 1371 0.01      
8 A1 1331 1261 11.83      
9 A1 1042 987 7.04      
10 A1 747 707 0.03      
11 A1 342 324 0.11      
12 A2 3138 2972 0.00      
13 A2 1511 1431 0.00      
14 A2 1022 968 0.00      
15 A2 628 595 0.00      
16 A2 156 148 0.00      
17 B1 3250 3078 3.72      
18 B1 3138 2972 41.36      
19 B1 1529 1448 21.09      
20 B1 1116 1057 0.03      
21 B1 1038 983 0.07      
22 B1 429 406 0.67      
23 B1 259 245 6.01      
24 B1 166 157 1.52      
25 B2 3185 3016 12.78      
26 B2 3065 2903 18.44      
27 B2 1520 1440 9.29      
28 B2 1432 1356 9.29      
29 B2 1251 1185 3.82      
30 B2 992 939 0.31      
31 B2 863 818 56.48      
32 B2 598 566 10.35      
33 B2 181 171 0.55      

Unscaled Zero Point Vibrational Energy (zpe) 24944.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23625.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.27602 0.12012 0.08795

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.246
C2 0.000 0.000 0.932
H3 -0.927 0.000 2.812
H4 0.927 0.000 2.812
C5 0.000 0.000 -0.383
C6 0.000 1.285 -1.173
C7 0.000 -1.285 -1.173
H8 0.000 2.157 -0.518
H9 0.000 -2.157 -0.518
H10 0.882 1.335 -1.821
H11 -0.882 1.335 -1.821
H12 -0.882 -1.335 -1.821
H13 0.882 -1.335 -1.821

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 H8 H9 H10 H11 H12 H13
C11.31391.08631.08632.62853.65253.65253.50663.50664.36994.36994.36994.3699
C21.31392.09612.09611.31462.46642.46642.59972.59973.18373.18373.18373.1837
H31.08632.09611.85453.32634.28854.28854.07494.07495.14924.82134.82135.1492
H41.08632.09611.85453.32634.28854.28854.07494.07494.82135.14925.14924.8213
C52.62851.31463.32633.32631.50891.50892.16172.16172.15092.15092.15092.1509
C63.65252.46644.28854.28851.50892.57051.09053.50441.09501.09502.83902.8390
C73.65252.46644.28854.28851.50892.57053.50441.09052.83902.83901.09501.0950
H83.50662.59974.07494.07492.16171.09053.50444.31491.77491.77493.82983.8298
H93.50662.59974.07494.07492.16173.50441.09054.31493.82983.82981.77491.7749
H104.36993.18375.14924.82132.15091.09502.83901.77493.82981.76313.19892.6691
H114.36993.18374.82135.14922.15091.09502.83901.77493.82981.76312.66913.1989
H124.36993.18374.82135.14922.15092.83901.09503.82981.77493.19892.66911.7631
H134.36993.18375.14924.82132.15092.83901.09503.82981.77492.66913.19891.7631

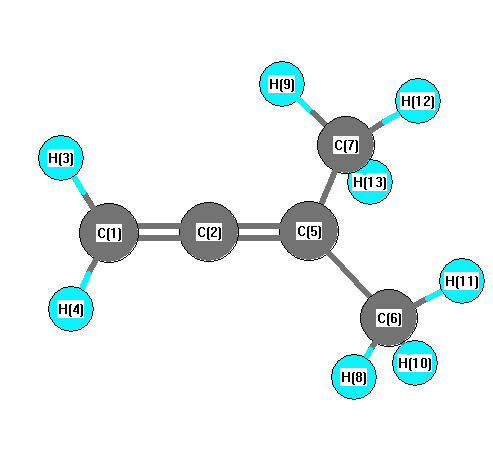
picture of 1,2-Butadiene, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C5 180.000 C2 C1 H3 121.399
C2 C1 H4 121.399 C2 C5 C6 121.593
C2 C5 C7 121.593 H3 C1 H4 117.201
C5 C6 H8 111.517 C5 C6 H10 110.379
C5 C6 H11 110.379 C5 C7 H9 111.517
C5 C7 H12 110.379 C5 C7 H13 110.379
C6 C5 C7 116.815 H8 C6 H10 108.606
H8 C6 H11 108.606 H9 C7 H12 108.606
H9 C7 H13 108.606 H10 C6 H11 107.228
H12 C7 H13 107.228
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability