return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Calicene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-307.542790
Energy at 298.15K-307.547143
HF Energy-306.341818
Nuclear repulsion energy299.340500
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3326 3150 11.01      
2 A1 3259 3087 15.14      
3 A1 3237 3066 5.40      
4 A1 1849 1751 568.11      
5 A1 1561 1478 60.69      
6 A1 1499 1420 36.10      
7 A1 1421 1345 101.35      
8 A1 1224 1159 31.54      
9 A1 1089 1031 13.98      
10 A1 1042 987 10.03      
11 A1 957 907 15.37      
12 A1 888 841 33.28      
13 A1 488 462 0.24      
14 A2 869 823 0.00      
15 A2 776 735 0.00      
16 A2 630 597 0.00      
17 A2 540 511 0.00      
18 A2 209 198 0.00      
19 B1 807 764 1.69      
20 B1 693 657 158.41      
21 B1 640 606 7.02      
22 B1 550 521 3.83      
23 B1 368 349 11.13      
24 B1 93 88 5.34      
25 B2 3287 3113 5.89      
26 B2 3251 3079 23.80      
27 B2 3228 3057 1.03      
28 B2 1569 1486 0.46      
29 B2 1364 1292 0.00      
30 B2 1243 1177 0.94      
31 B2 1114 1055 13.51      
32 B2 1099 1041 8.75      
33 B2 946 896 9.37      
34 B2 833 789 1.25      
35 B2 494 468 1.48      
36 B2 143 136 1.77      

Unscaled Zero Point Vibrational Energy (zpe) 23292.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22060.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.22511 0.04743 0.03918

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.049
C2 0.000 0.000 1.303
C3 0.000 0.668 2.572
C4 0.000 -0.668 2.572
C5 0.000 1.172 -0.907
C6 0.000 -1.172 -0.907
C7 0.000 0.726 -2.205
C8 0.000 -0.726 -2.205
H9 0.000 1.587 3.138
H10 0.000 -1.587 3.138
H11 0.000 2.200 -0.562
H12 0.000 -2.200 -0.562
H13 0.000 1.350 -3.093
H14 0.000 -1.350 -3.093

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.35222.70452.70451.45311.45312.27572.27573.56073.56072.25942.25943.33013.3301
C21.35221.43371.43372.50242.50243.58333.58332.42642.42642.88482.88484.59904.5990
C32.70451.43371.33693.51543.93614.77754.97661.07972.32603.48834.24875.70556.0136
C42.70451.43371.33693.93613.51544.97664.77752.32601.07974.24873.48836.01365.7055
C51.45312.50243.51543.93612.34481.37262.30004.06704.89751.08433.39032.19283.3376
C61.45312.50243.93613.51542.34482.30001.37264.89754.06703.39031.08433.33762.1928
C72.27573.58334.77754.97661.37262.30001.45265.41295.82332.20743.35641.08482.2581
C82.27573.58334.97664.77752.30001.37261.45265.82335.41293.35642.20742.25811.0848
H93.56072.42641.07972.32604.06704.89755.41295.82333.17503.75115.29556.23596.8891
H103.56072.42642.32601.07974.89754.06705.82335.41293.17505.29553.75116.88916.2359
H112.25942.88483.48834.24871.08433.39032.20743.35643.75115.29554.40062.66974.3600
H122.25942.88484.24873.48833.39031.08433.35642.20745.29553.75114.40064.36002.6697
H133.33014.59905.70556.01362.19283.33761.08482.25816.23596.88912.66974.36002.7002
H143.33014.59906.01365.70553.33762.19282.25811.08486.88916.23594.36002.66972.7002

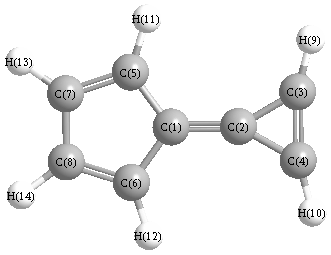
picture of Calicene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 152.209 C1 C2 C4 152.209
C1 C5 C7 107.250 C1 C5 H11 125.227
C1 C6 C8 107.250 C1 C6 H12 125.227
C2 C1 C5 126.215 C2 C1 C6 126.215
C2 C3 C4 62.209 C2 C3 H9 149.450
C2 C4 C3 62.209 C2 C4 H10 149.450
C3 C2 C4 55.581 C3 C4 H10 148.341
C4 C3 H9 148.341 C5 C1 C6 107.571
C5 C7 C8 108.965 C5 C7 H13 125.931
C6 C8 C7 108.965 C6 C8 H14 125.931
C7 C5 H11 127.523 C7 C8 H14 125.104
C8 C6 H12 127.523 C8 C7 H13 125.104
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability