Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.729 |
| C2 |
0.000 |
1.343 |
-0.417 |
| C3 |
0.000 |
-1.343 |
-0.417 |
| C4 |
0.797 |
2.410 |
-0.280 |
| C5 |
-0.797 |
-2.410 |
-0.280 |
| H6 |
-0.747 |
1.290 |
-1.205 |
| H7 |
0.747 |
-1.290 |
-1.205 |
| H8 |
0.706 |
3.255 |
-0.953 |
| H9 |
1.543 |
2.466 |
0.506 |
| H10 |
-0.706 |
-3.255 |
-0.953 |
| H11 |
-1.543 |
-2.466 |
0.506 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7649 | 1.7649 | 2.7317 | 2.7317 | 2.4417 | 2.4417 | 3.7317 | 2.9170 | 3.7317 | 2.9170 |
C2 | 1.7649 | | 2.6852 | 1.3394 | 3.8390 | 1.0872 | 2.8477 | 2.1083 | 2.1196 | 4.6827 | 4.2112 | C3 | 1.7649 | 2.6852 | | 3.8390 | 1.3394 | 2.8477 | 1.0872 | 4.6827 | 4.2112 | 2.1083 | 2.1196 | C4 | 2.7317 | 1.3394 | 3.8390 | | 5.0773 | 2.1203 | 3.8144 | 1.0846 | 1.0846 | 5.9002 | 5.4650 | C5 | 2.7317 | 3.8390 | 1.3394 | 5.0773 | | 3.8144 | 2.1203 | 5.9002 | 5.4650 | 1.0846 | 1.0846 | H6 | 2.4417 | 1.0872 | 2.8477 | 2.1203 | 3.8144 | | 2.9811 | 2.4571 | 3.0907 | 4.5524 | 4.2030 | H7 | 2.4417 | 2.8477 | 1.0872 | 3.8144 | 2.1203 | 2.9811 | | 4.5524 | 4.2030 | 2.4571 | 3.0907 | H8 | 3.7317 | 2.1083 | 4.6827 | 1.0846 | 5.9002 | 2.4571 | 4.5524 | | 1.8584 | 6.6621 | 6.3179 | H9 | 2.9170 | 2.1196 | 4.2112 | 1.0846 | 5.4650 | 3.0907 | 4.2030 | 1.8584 | | 6.3179 | 5.8169 | H10 | 3.7317 | 4.6827 | 2.1083 | 5.9002 | 1.0846 | 4.5524 | 2.4571 | 6.6621 | 6.3179 | | 1.8584 | H11 | 2.9170 | 4.2112 | 2.1196 | 5.4650 | 1.0846 | 4.2030 | 3.0907 | 6.3179 | 5.8169 | 1.8584 | |
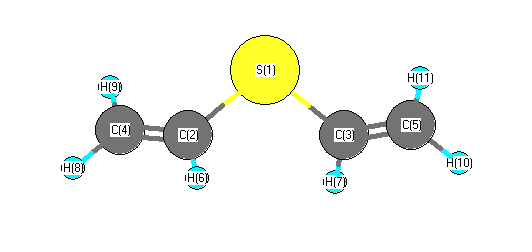 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.681 |
|
S1 |
C2 |
H6 |
115.706 |
| S1 |
C3 |
C5 |
122.681 |
|
S1 |
C3 |
H7 |
115.706 |
| C2 |
S1 |
C3 |
99.055 |
|
C2 |
C4 |
H8 |
120.502 |
| C2 |
C4 |
H9 |
121.593 |
|
C3 |
C5 |
H10 |
120.502 |
| C3 |
C5 |
H11 |
121.593 |
|
C4 |
C2 |
H6 |
121.443 |
| C5 |
C3 |
H7 |
121.443 |
|
H8 |
C4 |
H9 |
117.902 |
| H10 |
C5 |
H11 |
117.902 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -553.328312 |
| Energy at 298.15K | -553.333829 |
| HF Energy | -552.467578 |
| Nuclear repulsion energy | 202.746252 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3292 |
3118 |
5.16 |
|
|
|
| 2 |
A |
3290 |
3116 |
4.81 |
|
|
|
| 3 |
A |
3214 |
3044 |
6.61 |
|
|
|
| 4 |
A |
3201 |
3032 |
7.44 |
|
|
|
| 5 |
A |
3190 |
3021 |
0.59 |
|
|
|
| 6 |
A |
3184 |
3016 |
1.50 |
|
|
|
| 7 |
A |
1655 |
1568 |
24.50 |
|
|
|
| 8 |
A |
1644 |
1557 |
24.14 |
|
|
|
| 9 |
A |
1441 |
1364 |
3.80 |
|
|
|
| 10 |
A |
1437 |
1361 |
7.76 |
|
|
|
| 11 |
A |
1324 |
1254 |
1.94 |
|
|
|
| 12 |
A |
1318 |
1249 |
7.96 |
|
|
|
| 13 |
A |
1081 |
1024 |
8.41 |
|
|
|
| 14 |
A |
1056 |
1000 |
15.40 |
|
|
|
| 15 |
A |
1004 |
951 |
34.47 |
|
|
|
| 16 |
A |
989 |
936 |
42.11 |
|
|
|
| 17 |
A |
906 |
858 |
36.70 |
|
|
|
| 18 |
A |
863 |
818 |
43.17 |
|
|
|
| 19 |
A |
766 |
725 |
7.42 |
|
|
|
| 20 |
A |
720 |
682 |
4.95 |
|
|
|
| 21 |
A |
643 |
609 |
17.87 |
|
|
|
| 22 |
A |
607 |
575 |
16.95 |
|
|
|
| 23 |
A |
461 |
437 |
0.37 |
|
|
|
| 24 |
A |
379 |
359 |
0.38 |
|
|
|
| 25 |
A |
237 |
224 |
1.48 |
|
|
|
| 26 |
A |
149 |
141 |
4.50 |
|
|
|
| 27 |
A |
94 |
89 |
1.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19071.7 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18062.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.001 |
-1.007 |
-0.019 |
| C2 |
1.069 |
0.317 |
0.423 |
| C3 |
-1.547 |
-0.169 |
-0.020 |
| C4 |
2.205 |
0.583 |
-0.234 |
| C5 |
-1.741 |
1.154 |
-0.127 |
| H6 |
0.786 |
0.861 |
1.322 |
| H7 |
-2.388 |
-0.849 |
0.088 |
| H8 |
2.889 |
1.341 |
0.133 |
| H9 |
2.478 |
0.049 |
-1.137 |
| H10 |
-0.920 |
1.848 |
-0.265 |
| H11 |
-2.745 |
1.559 |
-0.086 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7585 | 1.7586 | 2.7279 | 2.7760 | 2.4302 | 2.3944 | 3.7265 | 2.9172 | 3.0096 | 3.7574 |
C2 | 1.7585 | | 2.6971 | 1.3395 | 2.9824 | 1.0880 | 3.6631 | 2.1084 | 2.1196 | 2.6027 | 4.0433 | C3 | 1.7586 | 2.6971 | | 3.8327 | 1.3408 | 2.8815 | 1.0868 | 4.6881 | 4.1827 | 2.1265 | 2.1034 | C4 | 2.7279 | 1.3395 | 3.8327 | | 3.9882 | 2.1242 | 4.8217 | 1.0847 | 1.0843 | 3.3717 | 5.0478 | C5 | 2.7760 | 2.9824 | 1.3408 | 3.9882 | | 2.9265 | 2.1156 | 4.6402 | 4.4764 | 1.0839 | 1.0840 | H6 | 2.4302 | 1.0880 | 2.8815 | 2.1242 | 2.9265 | | 3.8100 | 2.4628 | 3.0935 | 2.5301 | 3.8647 | H7 | 2.3944 | 3.6631 | 1.0868 | 4.8217 | 2.1156 | 3.8100 | | 5.7130 | 5.0973 | 3.0910 | 2.4403 | H8 | 3.7265 | 2.1084 | 4.6881 | 1.0847 | 4.6402 | 2.4628 | 5.7130 | | 1.8583 | 3.8628 | 5.6423 | H9 | 2.9172 | 2.1196 | 4.1827 | 1.0843 | 4.4764 | 3.0935 | 5.0973 | 1.8583 | | 3.9427 | 5.5378 | H10 | 3.0096 | 2.6027 | 2.1265 | 3.3717 | 1.0839 | 2.5301 | 3.0910 | 3.8628 | 3.9427 | | 1.8567 | H11 | 3.7574 | 4.0433 | 2.1034 | 5.0478 | 1.0840 | 3.8647 | 2.4403 | 5.6423 | 5.5378 | 1.8567 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.847 |
|
S1 |
C2 |
H6 |
115.214 |
| S1 |
C3 |
C5 |
126.659 |
|
S1 |
C3 |
H7 |
112.438 |
| C2 |
S1 |
C3 |
100.147 |
|
C2 |
C4 |
H8 |
120.491 |
| C2 |
C4 |
H9 |
121.611 |
|
C3 |
C5 |
H10 |
122.203 |
| C3 |
C5 |
H11 |
119.953 |
|
C4 |
C2 |
H6 |
121.765 |
| C5 |
C3 |
H7 |
120.901 |
|
H8 |
C4 |
H9 |
117.896 |
| H10 |
C5 |
H11 |
117.840 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
