return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1-Hexyne)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.936912
Energy at 298.15K-233.946116
HF Energy-233.009794
Nuclear repulsion energy210.593348
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3496 3311 53.33      
2 A' 3168 3000 31.79      
3 A' 3086 2923 54.46      
4 A' 3077 2914 21.59      
5 A' 3075 2912 3.29      
6 A' 3071 2908 14.41      
7 A' 2142 2029 1.20      
8 A' 1548 1466 11.01      
9 A' 1536 1454 2.28      
10 A' 1527 1446 0.69      
11 A' 1518 1437 1.41      
12 A' 1450 1373 5.97      
13 A' 1426 1350 1.01      
14 A' 1375 1303 2.38      
15 A' 1288 1220 1.05      
16 A' 1148 1087 4.38      
17 A' 1104 1046 0.16      
18 A' 1061 1004 1.37      
19 A' 964 913 0.29      
20 A' 927 878 3.75      
21 A' 563 534 51.55      
22 A' 472 447 2.42      
23 A' 392 371 0.06      
24 A' 257 243 0.37      
25 A' 111 105 0.03      
26 A" 3166 2999 52.08      
27 A" 3146 2979 29.69      
28 A" 3121 2956 0.02      
29 A" 3112 2948 4.69      
30 A" 1541 1460 10.39      
31 A" 1361 1289 0.06      
32 A" 1344 1273 0.79      
33 A" 1268 1201 0.07      
34 A" 1150 1089 0.13      
35 A" 951 901 0.13      
36 A" 804 762 0.86      
37 A" 735 696 2.83      
38 A" 547 518 53.15      
39 A" 287 272 0.99      
40 A" 251 238 0.05      
41 A" 113 107 0.00      
42 A" 86 82 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 31381.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29721.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.48654 0.04432 0.04188

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.236 2.625 0.000
H2 2.217 3.106 0.000
H3 0.693 2.972 0.883
H4 0.693 2.972 -0.883
C5 1.359 1.104 0.000
H6 1.929 0.779 0.878
H7 1.929 0.779 -0.878
C8 0.000 0.410 0.000
H9 -0.576 0.720 0.879
H10 -0.576 0.720 -0.879
C11 0.123 -1.119 0.000
H12 0.694 -1.442 0.878
H13 0.694 -1.442 -0.878
C14 -2.281 -2.303 0.000
H15 -3.239 -2.768 0.000
C16 -1.180 -1.783 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 H13 C14 H15 C16
C11.09261.09321.09321.52592.15802.15802.53642.77162.77163.90544.19524.19526.05427.00765.0260
H21.09261.76631.76632.17812.50312.50313.49043.77663.77664.71494.87534.87537.03488.01675.9527
H31.09321.76631.76612.17102.51723.07212.79742.58493.12804.22374.41374.75216.12007.01375.1861
H41.09321.76631.76612.17103.07212.51722.79743.12802.58494.22374.75214.41376.12007.01375.1861
C51.52592.17812.17102.17101.09631.09631.52582.15932.15932.54292.77352.77354.98566.01083.8440
H62.15802.50312.51723.07211.09631.75632.15182.50613.06042.76342.54153.08935.29176.32974.1234
H72.15802.50313.07212.51721.09631.75632.15183.06042.50612.76343.08932.54155.29176.32974.1234
C82.53643.49042.79742.79741.52582.15182.15181.09541.09541.53352.16332.16333.54454.53752.4898
H92.77163.77662.58493.12802.15932.50613.06041.09541.75702.15472.50723.06143.58054.47572.7205
H102.77163.77663.12802.58492.15933.06042.50611.09541.75702.15473.06142.50723.58054.47572.7205
C113.90544.71494.22374.22372.54292.76342.76341.53352.15472.15471.09591.09592.68033.74461.4624
H124.19524.87534.41374.75212.77352.54153.08932.16332.50723.06141.09591.75633.21944.24202.0972
H134.19524.87534.75214.41372.77353.08932.54152.16333.06142.50721.09591.75633.21944.24202.0972
C146.05427.03486.12006.12004.98565.29175.29173.54453.58053.58052.68033.21943.21941.06431.2182
H157.00768.01677.01377.01376.01086.32976.32974.53754.47574.47573.74464.24204.24201.06432.2825
C165.02605.95275.18615.18613.84404.12344.12342.48982.72052.72051.46242.09722.09721.21822.2825

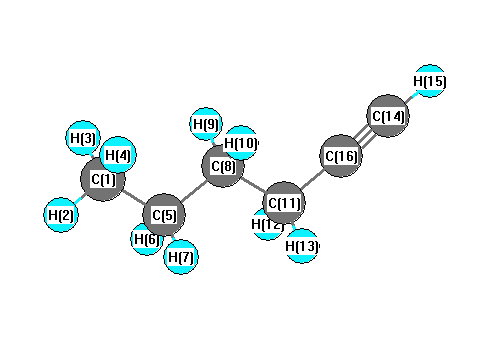
picture of 1-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.683 C1 C5 H7 109.683
C1 C5 C8 112.436 H2 C1 H3 107.813
H2 C1 H4 107.813 H2 C1 C5 111.500
H3 C1 H4 107.756 H3 C1 C5 110.900
H4 C1 C5 110.900 C5 C8 H9 109.838
C5 C8 H10 109.838 C5 C8 C11 112.445
H6 C5 H7 106.453 H6 C5 C8 109.202
H7 C5 C8 109.202 C8 C11 H12 109.598
C8 C11 H13 109.598 C8 C11 C16 112.389
H9 C8 H10 106.640 H9 C8 C11 108.952
H10 C8 C11 108.952 C11 C16 C14 178.285
H12 C11 H13 106.517 H12 C11 C16 109.282
H13 C11 C16 109.282 H15 C14 C16 179.346
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability