return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NS (4-Methylthiazole)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-607.447905
Energy at 298.15K-607.453926
HF Energy-606.387926
Nuclear repulsion energy273.853024
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3281 3108 1.31      
2 A 3259 3086 0.29      
3 A 3184 3016 12.20      
4 A 3169 3001 10.66      
5 A 3086 2922 19.42      
6 A 1566 1483 14.77      
7 A 1518 1438 17.53      
8 A 1511 1431 9.90      
9 A 1452 1375 16.96      
10 A 1434 1358 29.60      
11 A 1369 1297 15.43      
12 A 1272 1204 0.78      
13 A 1192 1129 2.60      
14 A 1069 1012 1.43      
15 A 1028 974 7.16      
16 A 963 912 20.10      
17 A 927 878 28.12      
18 A 871 825 12.02      
19 A 774 733 58.51      
20 A 695 658 0.91      
21 A 693 656 6.97      
22 A 632 598 1.39      
23 A 567 537 1.25      
24 A 474 449 0.49      
25 A 333 315 2.78      
26 A 228 216 5.14      
27 A 107 101 0.49      

Unscaled Zero Point Vibrational Energy (zpe) 18325.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17356.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.24491 0.08416 0.06338

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 1.566 2.015 0.000
C2 -0.950 -0.032 -0.000
C3 0.913 1.150 0.000
H4 -2.868 0.304 0.882
H5 -2.735 -1.229 -0.002
H6 -2.868 0.307 -0.881
C7 -2.440 -0.178 0.000
S8 1.561 -0.445 0.000
N9 -0.398 1.225 -0.000
H10 -0.224 -2.126 -0.000
C11 -0.033 -1.061 -0.000

Atom - Atom Distances (Å)
  H1 C2 C3 H4 H5 H6 C7 S8 N9 H10 C11
H13.24341.08374.83335.38624.83224.56612.45992.11624.51083.4666
C23.24342.20662.13712.14822.13701.49632.54501.37342.21651.3786
C31.08372.20663.97354.35443.97283.60591.72131.31313.46722.4047
H44.83332.13713.97351.77401.76281.09254.57722.77993.69763.2675
H55.38622.14824.35441.77401.77411.09154.36623.38852.66632.7066
H64.83222.13703.97281.76281.77411.09244.57772.77873.69933.2684
C74.56611.49633.60591.09251.09151.09244.00942.47742.95052.5635
S82.45992.54501.72134.57724.36624.57774.00942.57432.45151.7088
N92.11621.37341.31312.77993.38852.77872.47742.57433.35582.3154
H104.51082.21653.46723.69762.66633.69932.95052.45153.35581.0817
C113.46661.37862.40473.26752.70663.26842.56351.70882.31541.0817

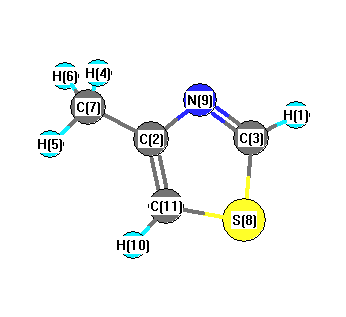
picture of 4-Methylthiazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C3 S8 120.869 H1 C3 N9 123.719
C2 C7 H4 110.313 C2 C7 H5 111.262
C2 C7 H6 110.307 C2 N9 C3 110.417
C2 C11 S8 110.582 C2 C11 H10 128.137
C3 S8 C11 89.025 H4 C7 H5 108.641
H4 C7 H6 107.573 H5 C7 H6 108.644
C7 C2 N9 119.312 C7 C2 C11 126.124
S8 C3 N9 115.412 S8 C11 H10 121.281
N9 C2 C11 114.564
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability