return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,4-Hexadiene, (Z)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.946356
Energy at 298.15K-233.955971
HF Energy-233.023323
Nuclear repulsion energy221.098406
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3286 3113 10.75      
2 A 3201 3032 37.75      
3 A 3192 3023 12.63      
4 A 3188 3019 9.88      
5 A 3174 3006 21.13      
6 A 3172 3005 0.27      
7 A 3144 2978 15.07      
8 A 3112 2947 14.33      
9 A 3075 2913 19.68      
10 A 3072 2909 23.08      
11 A 1723 1632 1.88      
12 A 1702 1612 8.17      
13 A 1543 1461 10.22      
14 A 1524 1443 3.56      
15 A 1518 1437 16.22      
16 A 1464 1386 6.52      
17 A 1455 1378 6.02      
18 A 1427 1351 3.89      
19 A 1371 1298 1.16      
20 A 1333 1262 2.11      
21 A 1301 1232 0.87      
22 A 1255 1189 1.32      
23 A 1142 1081 1.31      
24 A 1107 1049 7.18      
25 A 1076 1019 1.44      
26 A 1029 974 23.88      
27 A 1019 965 1.85      
28 A 988 936 1.34      
29 A 967 916 0.28      
30 A 940 890 4.21      
31 A 902 854 37.54      
32 A 884 837 1.83      
33 A 739 700 35.53      
34 A 629 596 8.52      
35 A 554 525 14.14      
36 A 534 506 6.64      
37 A 395 374 0.45      
38 A 301 285 0.04      
39 A 214 203 0.25      
40 A 125 118 0.02      
41 A 103 98 0.20      
42 A 44 42 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 31460.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29796.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18653 0.07848 0.06918

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.907 0.787 -0.569
H2 2.816 0.450 -1.078
H3 1.114 0.892 -1.310
H4 2.118 1.782 -0.167
C5 1.527 -0.154 0.536
H6 2.212 -0.202 1.382
C7 0.432 -0.931 0.571
H8 0.266 -1.543 1.456
C9 -0.622 -1.002 -0.499
H10 -0.171 -0.894 -1.492
H11 -1.071 -2.002 -0.493
C12 -1.728 0.018 -0.355
H13 -2.582 -0.119 -1.018
C14 -1.724 1.057 0.488
H15 -2.558 1.749 0.522
H16 -0.895 1.233 1.166

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 H10 H11 C12 H13 C14 H15 H16
C11.09481.09021.09381.50012.20852.53483.49583.09772.82744.07993.72124.60153.79094.69563.3261
H21.09481.77351.75792.15202.61533.21084.11033.77623.30144.63224.62125.42854.84035.75524.4071
H31.09021.77351.76222.16193.10652.70733.78192.69442.20833.71723.12333.84363.36404.19223.2074
H41.09381.75791.76222.14242.51883.27814.13733.91953.76244.95874.23525.14093.96394.72623.3404
C51.50012.15202.16192.14241.08941.34332.08952.53092.74683.34993.37904.39353.46884.50612.8615
H62.20852.61533.10652.51881.08942.08742.36423.49343.79694.18684.31145.36214.22755.22473.4298
C72.53483.21082.70733.27811.34332.08741.08871.50292.14982.13002.53433.50282.93314.01522.6075
H83.49584.11033.78194.13732.08952.36421.08872.21383.05022.40773.11334.03273.41364.43663.0233
C93.09773.77622.69443.91952.53093.49341.50292.21381.09661.09651.51112.21212.53473.51512.8000
H102.82743.30142.20833.76242.74683.79692.14983.05021.09661.74192.13302.57723.18454.09173.4811
H114.07994.63223.71724.95873.34994.18682.13002.40771.09651.74192.12872.47143.27824.16103.6402
C123.72124.62123.12334.23523.37904.31142.53433.11331.51112.13302.12871.09001.33832.11072.1176
H134.60155.42853.84365.14094.39355.36213.50284.03272.21212.57722.47141.09002.09502.42103.0731
C143.79094.84033.36403.96393.46884.22752.93313.41362.53473.18453.27821.33832.09501.08461.0844
H154.69565.75524.19224.72624.50615.22474.01524.43663.51514.09174.16102.11072.42101.08461.8558
H163.32614.40713.20743.34042.86153.42982.60753.02332.80003.48113.64022.11763.07311.08441.8558

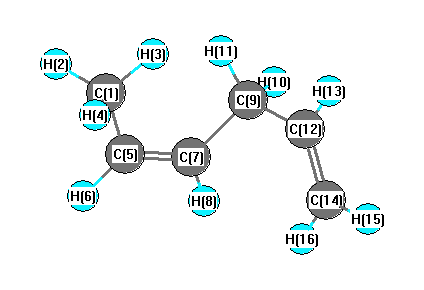
picture of 1,4-Hexadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 116.145 C1 C5 C7 126.036
H2 C1 H3 108.513 H2 C1 H4 106.871
H2 C1 C5 111.094 H3 C1 H4 107.580
H3 C1 C5 112.186 H4 C1 C5 110.387
C5 C7 H8 118.069 C5 C7 C9 125.465
H6 C5 C7 117.819 C7 C9 H10 110.615
C7 C9 H11 109.057 C7 C9 C12 114.453
H8 C7 C9 116.439 C9 C12 H13 115.567
C9 C12 C14 125.527 H10 C9 H11 105.173
H10 C9 C12 108.717 H11 C9 C12 108.396
C12 C14 H15 120.831 C12 C14 H16 121.512
H13 C12 C14 118.898 H15 C14 H16 117.655
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability