return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Butadiene, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-194.755922
Energy at 298.15K-194.763524
HF Energy-193.991445
Nuclear repulsion energy160.263614
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3287 3113 12.26      
2 A' 3275 3102 12.13      
3 A' 3195 3026 4.18      
4 A' 3182 3014 14.96      
5 A' 3180 3012 17.44      
6 A' 3178 3010 0.31      
7 A' 3076 2914 16.49      
8 A' 1711 1620 1.71      
9 A' 1659 1572 13.79      
10 A' 1537 1456 9.37      
11 A' 1482 1403 1.23      
12 A' 1456 1379 5.55      
13 A' 1437 1361 9.88      
14 A' 1356 1284 0.49      
15 A' 1335 1265 0.24      
16 A' 1101 1043 4.52      
17 A' 1024 970 0.48      
18 A' 984 932 0.37      
19 A' 808 765 0.20      
20 A' 536 508 0.20      
21 A' 426 404 1.09      
22 A' 286 271 0.67      
23 A" 3152 2985 15.34      
24 A" 1518 1438 10.80      
25 A" 1074 1018 0.24      
26 A" 1001 948 27.23      
27 A" 884 837 47.63      
28 A" 863 818 42.78      
29 A" 770 729 0.93      
30 A" 632 598 0.32      
31 A" 383 363 10.21      
32 A" 217 205 0.42      
33 A" 144 137 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 25075.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23748.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.28490 0.13914 0.09513

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.589 1.737 0.000
C2 0.000 0.525 0.000
C3 -0.830 -0.685 0.000
C4 -0.364 -1.947 0.000
C5 1.496 0.373 0.000
H6 -1.670 1.838 0.000
H7 -0.008 2.653 0.000
H8 -1.908 -0.526 0.000
H9 0.697 -2.173 0.000
H10 -1.044 -2.791 0.000
H11 1.988 1.347 0.000
H12 1.832 -0.182 0.881
H13 1.832 -0.182 -0.881

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.34762.43383.69022.49181.08571.08522.61884.11564.55052.60653.21243.2124
C21.34761.46702.49791.50382.12482.12862.17782.78603.47622.15122.15222.1522
C32.43381.46701.34472.55512.65963.43801.08962.13152.11683.47382.84852.8485
C43.69022.49791.34472.97294.00384.61362.09791.08451.08444.04672.95162.9516
C52.49181.50382.55512.97293.48902.73213.52042.66804.05731.09111.09391.0939
H61.08572.12482.65964.00383.48901.85112.37604.65734.67143.69114.13804.1380
H71.08522.12863.43804.61362.73211.85113.70344.87735.54202.38583.49303.4930
H82.61882.17781.08962.09793.52042.37603.70343.08142.42414.32233.85743.8574
H94.11562.78602.13151.08452.66804.65734.87733.08141.84733.74882.45532.4553
H104.55053.47622.11681.08444.05734.67145.54202.42411.84735.12963.98193.9819
H112.60652.15123.47384.04671.09113.69112.38584.32233.74885.12961.77111.7711
H123.21242.15222.84852.95161.09394.13803.49303.85742.45533.98191.77111.7619
H133.21242.15222.84852.95161.09394.13803.49303.85742.45533.98191.77111.7619

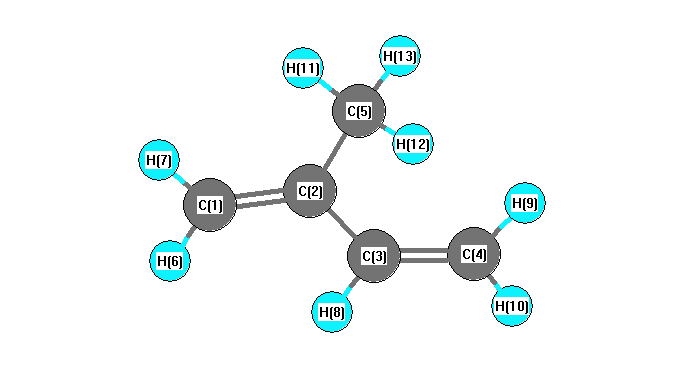
picture of 1,3-Butadiene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.632 C1 C2 C5 121.728
C2 C1 H6 121.288 C2 C1 H7 121.704
C2 C3 C4 125.289 C2 C3 H8 116.042
C2 C5 H11 110.993 C2 C5 H12 110.902
C2 C5 H13 110.902 C3 C2 C5 118.640
C3 C4 H9 122.311 C3 C4 H10 120.879
C4 C3 H8 118.670 H6 C1 H7 117.008
H9 C4 H10 116.810 H11 C5 H12 108.309
H11 C5 H13 108.309 H12 C5 H13 107.292
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability