return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH(CH3)CH3 (Propanal, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-231.868890
Energy at 298.15K-231.877415
HF Energy-231.032620
Nuclear repulsion energy175.632626
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3172 3004 23.37      
2 A' 3165 2998 46.60      
3 A' 3120 2955 4.68      
4 A' 3075 2912 15.40      
5 A' 2917 2762 112.16      
6 A' 1785 1691 129.42      
7 A' 1551 1469 23.45      
8 A' 1544 1462 8.81      
9 A' 1458 1380 5.56      
10 A' 1437 1361 3.30      
11 A' 1342 1271 0.34      
12 A' 1214 1150 0.03      
13 A' 1206 1142 8.19      
14 A' 958 907 16.38      
15 A' 866 821 22.34      
16 A' 556 527 5.73      
17 A' 360 341 6.60      
18 A' 338 320 0.19      
19 A' 245 232 0.82      
20 A" 3171 3004 12.73      
21 A" 3160 2993 3.13      
22 A" 3073 2911 21.82      
23 A" 1531 1450 3.18      
24 A" 1524 1443 1.35      
25 A" 1435 1359 8.77      
26 A" 1373 1300 0.37      
27 A" 1167 1105 1.02      
28 A" 1014 960 0.80      
29 A" 970 918 0.29      
30 A" 949 898 1.55      
31 A" 341 323 0.65      
32 A" 217 205 0.03      
33 A" 60 56 5.22      

Unscaled Zero Point Vibrational Energy (zpe) 25145.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23814.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.25796 0.12428 0.09402

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.226 0.589 2.167
H2 -0.226 0.589 -2.167
H3 -1.700 0.113 -1.306
H4 -1.700 0.113 1.306
H5 -1.164 1.796 1.275
H6 -1.164 1.796 -1.275
C7 -0.817 0.759 -1.264
C8 -0.817 0.759 1.264
H9 0.909 1.094 0.000
C10 0.000 0.484 0.000
C11 0.446 -0.957 0.000
H12 -0.388 -1.697 0.000
O13 1.599 -1.333 0.000

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 H9 C10 C11 H12 O13
H14.33343.80241.77161.76893.76563.48511.09262.49732.18102.74543.15403.4237
H24.33341.77163.80243.76561.76891.09263.48512.49732.18102.74543.15403.4237
H33.80241.77162.61223.12731.76591.09442.79283.07742.17532.73012.58903.8312
H41.77163.80242.61221.76593.12732.79281.09443.07742.17532.73012.58903.8312
H51.76893.76563.12731.76592.55012.76391.09272.53252.16783.43403.79804.3643
H63.76561.76891.76593.12732.55011.09272.76392.53252.16783.43403.79804.3643
C73.48511.09261.09442.79282.76391.09272.52732.16511.52982.47742.79533.4370
C81.09263.48512.79281.09441.09272.76392.52732.16511.52982.47742.79533.4370
H92.49732.49733.07743.07742.53252.53252.16512.16511.09422.10223.07702.5231
C102.18102.18102.17532.17532.16782.16781.52981.52981.09421.50852.21532.4207
C112.74542.74542.73012.73013.43403.43402.47742.47742.10221.50851.11451.2130
H123.15403.15402.58902.58903.79803.79802.79532.79533.07702.21531.11452.0197
O133.42373.42373.83123.83124.36434.36433.43703.43702.52312.42071.21302.0197

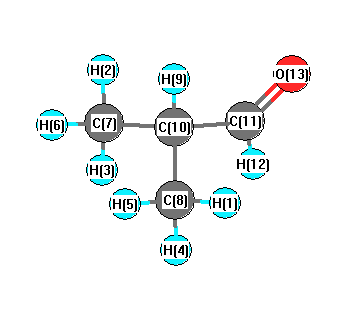
picture of Propanal, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C8 H4 108.204 H1 C8 H5 108.080
H1 C8 C10 111.449 H2 C7 H3 108.204
H2 C7 H6 108.080 H2 C7 C10 111.449
H3 C7 H6 107.690 H3 C7 C10 110.891
H4 C8 H5 107.690 H4 C8 C10 110.891
H5 C8 C10 110.394 H6 C7 C10 110.394
C7 C10 C8 111.383 C7 C10 H9 110.093
C7 C10 C11 109.245 C8 C10 H9 110.093
C8 C10 C11 109.245 H9 C10 C11 106.660
C10 C11 H12 114.410 C10 C11 O13 125.255
H12 C11 O13 120.334
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability