Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -115.644131 |
| Energy at 298.15K | -115.645862 |
| HF Energy | -115.212497 |
| Nuclear repulsion energy | 57.757472 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3336 |
3160 |
0.29 |
|
|
|
| 2 |
A' |
3127 |
2961 |
51.61 |
|
|
|
| 3 |
A' |
1633 |
1547 |
0.42 |
|
|
|
| 4 |
A' |
1262 |
1195 |
8.80 |
|
|
|
| 5 |
A' |
1033 |
978 |
18.59 |
|
|
|
| 6 |
A' |
935 |
886 |
9.07 |
|
|
|
| 7 |
A' |
596 |
564 |
77.32 |
|
|
|
| 8 |
A" |
5052 |
4785 |
99578.45 |
|
|
|
| 9 |
A" |
3041 |
2880 |
3747.66 |
|
|
|
| 10 |
A" |
1018 |
964 |
20.21 |
|
|
|
| 11 |
A" |
984 |
932 |
0.15 |
|
|
|
| 12 |
A" |
855 |
810 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11436.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 10831.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.037 |
0.891 |
0.000 |
| H2 |
0.750 |
1.646 |
0.000 |
| C3 |
-0.037 |
-0.419 |
0.662 |
| C4 |
-0.037 |
-0.419 |
-0.662 |
| H5 |
-0.040 |
-0.983 |
1.581 |
| H6 |
-0.040 |
-0.983 |
-1.581 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.0909 | 1.4672 | 1.4672 | 2.4512 | 2.4512 |
H2 | 1.0909 | | 2.3069 | 2.3069 | 3.1677 | 3.1677 | C3 | 1.4672 | 2.3069 | | 1.3232 | 1.0782 | 2.3120 | C4 | 1.4672 | 2.3069 | 1.3232 | | 2.3120 | 1.0782 | H5 | 2.4512 | 3.1677 | 1.0782 | 2.3120 | | 3.1611 | H6 | 2.4512 | 3.1677 | 2.3120 | 1.0782 | 3.1611 | |
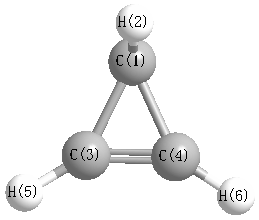 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.196 |
|
C1 |
C3 |
H5 |
148.344 |
| C1 |
C4 |
C3 |
63.196 |
|
C1 |
C4 |
H6 |
148.344 |
| H2 |
C1 |
C3 |
128.188 |
|
H2 |
C1 |
C4 |
128.188 |
| C3 |
C1 |
C4 |
53.608 |
|
C3 |
C4 |
H6 |
148.459 |
| C4 |
C3 |
H5 |
148.459 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
