Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.489505 |
| Energy at 298.15K | -169.493513 |
| HF Energy | -168.935548 |
| Nuclear repulsion energy | 70.743650 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3874 |
3669 |
64.17 |
|
|
|
| 2 |
A' |
3579 |
3390 |
5.60 |
|
|
|
| 3 |
A' |
3088 |
2925 |
63.22 |
|
|
|
| 4 |
A' |
1765 |
1671 |
150.04 |
|
|
|
| 5 |
A' |
1463 |
1385 |
11.64 |
|
|
|
| 6 |
A' |
1371 |
1298 |
173.19 |
|
|
|
| 7 |
A' |
1225 |
1160 |
69.94 |
|
|
|
| 8 |
A' |
1089 |
1032 |
202.27 |
|
|
|
| 9 |
A' |
626 |
593 |
1.20 |
|
|
|
| 10 |
A" |
1051 |
995 |
5.93 |
|
|
|
| 11 |
A" |
853 |
808 |
73.07 |
|
|
|
| 12 |
A" |
393 |
372 |
87.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10188.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 9649.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.376 |
0.000 |
| O2 |
-1.006 |
-0.528 |
0.000 |
| N3 |
1.202 |
-0.024 |
0.000 |
| H4 |
-0.334 |
1.422 |
0.000 |
| H5 |
-1.848 |
-0.067 |
0.000 |
| H6 |
1.819 |
0.782 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3530 | 1.2665 | 1.0974 | 1.9000 | 1.8639 |
O2 | 1.3530 | | 2.2646 | 2.0628 | 0.9597 | 3.1144 | N3 | 1.2665 | 2.2646 | | 2.1088 | 3.0495 | 1.0154 | H4 | 1.0974 | 2.0628 | 2.1088 | | 2.1233 | 2.2460 | H5 | 1.9000 | 0.9597 | 3.0495 | 2.1233 | | 3.7638 | H6 | 1.8639 | 3.1144 | 1.0154 | 2.2460 | 3.7638 | |
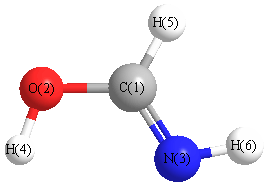 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.302 |
|
C1 |
N3 |
H6 |
109.045 |
| O2 |
C1 |
N3 |
119.615 |
|
O2 |
C1 |
H4 |
114.262 |
| N3 |
C1 |
H4 |
126.123 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.501070 |
| Energy at 298.15K | -169.505258 |
| HF Energy | -168.947376 |
| Nuclear repulsion energy | 71.128740 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3768 |
3568 |
45.71 |
|
|
|
| 2 |
A' |
3588 |
3399 |
6.23 |
|
|
|
| 3 |
A' |
3177 |
3009 |
32.02 |
|
|
|
| 4 |
A' |
1741 |
1649 |
231.48 |
|
|
|
| 5 |
A' |
1428 |
1352 |
26.60 |
|
|
|
| 6 |
A' |
1407 |
1333 |
1.88 |
|
|
|
| 7 |
A' |
1230 |
1164 |
114.49 |
|
|
|
| 8 |
A' |
1097 |
1039 |
183.58 |
|
|
|
| 9 |
A' |
595 |
563 |
56.25 |
|
|
|
| 10 |
A" |
1066 |
1010 |
0.90 |
|
|
|
| 11 |
A" |
852 |
807 |
34.25 |
|
|
|
| 12 |
A" |
661 |
626 |
219.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10304.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 9759.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
-1.113 |
-0.332 |
0.000 |
| N3 |
1.159 |
-0.101 |
0.000 |
| H4 |
-0.262 |
1.479 |
0.000 |
| H5 |
-0.817 |
-1.253 |
0.000 |
| H6 |
1.872 |
0.620 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3430 | 1.2702 | 1.0912 | 1.8612 | 1.8830 |
O2 | 1.3430 | | 2.2835 | 2.0006 | 0.9673 | 3.1336 | N3 | 1.2702 | 2.2835 | | 2.1247 | 2.2868 | 1.0146 | H4 | 1.0912 | 2.0006 | 2.1247 | | 2.7873 | 2.3007 | H5 | 1.8612 | 0.9673 | 2.2868 | 2.7873 | | 3.2772 | H6 | 1.8830 | 3.1336 | 1.0146 | 2.3007 | 3.2772 | |
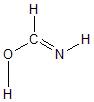 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.194 |
|
C1 |
N3 |
H6 |
110.507 |
| O2 |
C1 |
N3 |
121.784 |
|
O2 |
C1 |
H4 |
110.121 |
| N3 |
C1 |
H4 |
128.095 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.494907 |
| Energy at 298.15K | -169.499022 |
| HF Energy | -168.940813 |
| Nuclear repulsion energy | 70.626008 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3800 |
3599 |
28.43 |
|
|
|
| 2 |
A' |
3497 |
3312 |
7.30 |
|
|
|
| 3 |
A' |
3243 |
3071 |
12.18 |
|
|
|
| 4 |
A' |
1731 |
1639 |
231.80 |
|
|
|
| 5 |
A' |
1440 |
1364 |
0.41 |
|
|
|
| 6 |
A' |
1385 |
1312 |
26.19 |
|
|
|
| 7 |
A' |
1155 |
1094 |
282.90 |
|
|
|
| 8 |
A' |
1104 |
1046 |
46.85 |
|
|
|
| 9 |
A' |
599 |
567 |
37.77 |
|
|
|
| 10 |
A" |
1101 |
1043 |
87.31 |
|
|
|
| 11 |
A" |
855 |
810 |
53.89 |
|
|
|
| 12 |
A" |
537 |
509 |
93.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10223.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 9682.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.437 |
0.000 |
| O2 |
-1.105 |
-0.349 |
0.000 |
| N3 |
1.230 |
0.122 |
0.000 |
| H4 |
-0.298 |
1.482 |
0.000 |
| H5 |
-0.841 |
-1.277 |
0.000 |
| H6 |
1.370 |
-0.889 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3564 | 1.2695 | 1.0863 | 1.9098 | 1.9070 |
O2 | 1.3564 | | 2.3820 | 2.0010 | 0.9646 | 2.5332 | N3 | 1.2695 | 2.3820 | | 2.0452 | 2.4998 | 1.0215 | H4 | 1.0863 | 2.0010 | 2.0452 | | 2.8120 | 2.8992 | H5 | 1.9098 | 0.9646 | 2.4998 | 2.8120 | | 2.2451 | H6 | 1.9070 | 2.5332 | 1.0215 | 2.8992 | 2.2451 | |
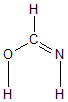 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.584 |
|
C1 |
N3 |
H6 |
112.244 |
| O2 |
C1 |
N3 |
130.198 |
|
O2 |
C1 |
H4 |
109.508 |
| N3 |
C1 |
H4 |
120.294 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
