return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6S (Thiophene, 2,5-dihydro-)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-550.248320
Energy at 298.15K-550.255120
HF Energy-549.749211
Nuclear repulsion energy213.866085
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3239 3071 10.09      
2 A1 3103 2942 0.37      
3 A1 1675 1588 1.33      
4 A1 1581 1499 8.70      
5 A1 1329 1260 0.34      
6 A1 1206 1143 0.58      
7 A1 931 883 0.15      
8 A1 715 678 3.11      
9 A1 534 506 0.06      
10 A2 3148 2984 0.00      
11 A2 1215 1152 0.00      
12 A2 1015 962 0.00      
13 A2 967 917 0.00      
14 A2 376 356 0.00      
15 B1 3150 2986 15.34      
16 B1 1201 1138 8.60      
17 B1 956 906 0.18      
18 B1 707 670 57.52      
19 B1 84 80 4.63      
20 B2 3211 3044 1.63      
21 B2 3104 2942 36.29      
22 B2 1584 1502 0.00      
23 B2 1410 1336 6.08      
24 B2 1302 1235 3.73      
25 B2 957 908 6.37      
26 B2 843 799 0.56      
27 B2 639 606 0.36      

Unscaled Zero Point Vibrational Energy (zpe) 20089.3 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 19044.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.21735 0.15590 0.09403

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.260
C2 0.000 1.361 0.005
C3 0.000 -1.361 0.005
C4 0.000 0.670 -1.354
C5 0.000 -0.670 -1.354
H6 -0.892 1.982 0.136
H7 0.892 1.982 0.136
H8 0.892 -1.982 0.136
H9 -0.892 -1.982 0.136
H10 0.000 1.274 -2.256
H11 0.000 -1.274 -2.256

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
S11.85151.85152.69872.69872.44682.44682.44682.44683.73993.7399
C21.85152.72251.52472.44411.09441.09443.46273.46272.26293.4722
C31.85152.72252.44411.52473.46273.46271.09441.09443.47222.2629
C42.69871.52472.44411.34032.17652.17653.17033.17031.08542.1430
C52.69872.44411.52471.34033.17033.17032.17652.17652.14301.0854
H62.44681.09443.46272.17653.17031.78314.34683.96422.64974.1375
H72.44681.09443.46272.17653.17031.78313.96424.34682.64974.1375
H82.44683.46271.09443.17032.17654.34683.96421.78314.13752.6497
H92.44683.46271.09443.17032.17653.96424.34681.78314.13752.6497
H103.73992.26293.47221.08542.14302.64972.64974.13754.13752.5474
H113.73993.47222.26292.14301.08544.13754.13752.64972.64972.5474

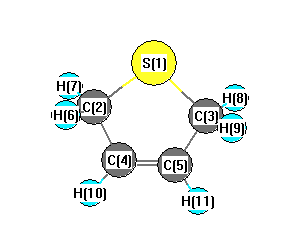
picture of Thiophene, 2,5-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C4 105.725 S1 C2 H6 109.622
S1 C2 H7 109.622 S1 C3 C5 105.725
S1 C3 H8 109.622 S1 C3 H9 109.622
C2 S1 C3 94.645 C2 C4 C5 116.953
C2 C4 H10 119.266 C3 C5 C4 116.953
C3 C5 H11 119.266 C4 C2 H6 111.350
C4 C2 H7 111.350 C4 C5 H11 123.782
C5 C3 H8 111.350 C5 C3 H9 111.350
C5 C4 H10 123.782 H6 C2 H7 109.113
H8 C3 H9 109.113
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability