return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (1,3-Cyclopentadiene)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-193.485725
Energy at 298.15K-193.491879
HF Energy-192.806243
Nuclear repulsion energy155.706799
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3283 3113 3.17      
2 A1 3258 3089 6.07      
3 A1 3080 2919 5.50      
4 A1 1554 1473 1.74      
5 A1 1417 1343 0.05      
6 A1 1395 1323 21.88      
7 A1 1122 1064 0.16      
8 A1 1037 983 0.04      
9 A1 947 898 10.62      
10 A1 804 762 0.02      
11 A2 1114 1056 0.00      
12 A2 928 879 0.00      
13 A2 703 667 0.00      
14 A2 507 481 0.00      
15 B1 3131 2969 4.88      
16 B1 938 889 1.19      
17 B1 908 861 24.21      
18 B1 681 645 59.79      
19 B1 325 308 9.18      
20 B2 3276 3105 15.73      
21 B2 3249 3080 2.20      
22 B2 1626 1541 0.25      
23 B2 1321 1253 1.52      
24 B2 1260 1194 0.92      
25 B2 1101 1044 1.99      
26 B2 989 938 9.82      
27 B2 807 765 4.99      

Unscaled Zero Point Vibrational Energy (zpe) 20380.2 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 19320.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.27837 0.27254 0.14137

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.886 0.000 1.882
H2 -0.886 0.000 1.882
H3 0.000 2.225 0.615
H4 0.000 -2.225 0.615
H5 0.000 1.360 -1.896
H6 0.000 -1.360 -1.896
C7 0.000 1.183 0.289
C8 0.000 -1.183 0.289
C9 0.000 0.000 1.220
C10 0.000 0.736 -1.000
C11 0.000 -0.736 -1.000

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 C9 C10 C11
H11.77262.70972.70974.11224.11222.17322.17321.10643.10363.1036
H21.77262.70972.70974.11224.11222.17322.17321.10643.10363.1036
H32.70972.70974.44972.65534.37681.09183.42322.30572.19643.3720
H42.70972.70974.44974.37682.65533.42321.09182.30573.37202.1964
H54.11224.11222.65534.37682.72062.19213.35283.39981.09252.2795
H64.11224.11224.37682.65532.72063.35282.19213.39982.27951.0925
C72.17322.17321.09183.42322.19213.35282.36561.50521.36412.3111
C82.17322.17323.42321.09183.35282.19212.36561.50522.31111.3641
C91.10641.10642.30572.30573.39983.39981.50521.50522.33832.3383
C103.10363.10362.19643.37201.09252.27951.36412.31112.33831.4713
C113.10363.10363.37202.19642.27951.09252.31111.36412.33831.4713

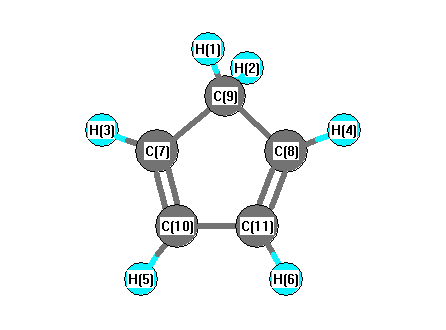
picture of 1,3-Cyclopentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C9 H2 106.463 H1 C9 C7 111.728
H1 C9 C8 111.728 H2 C9 C7 111.728
H2 C9 C8 111.728 H3 C7 C9 124.441
H3 C7 C10 126.490 H4 C8 C9 124.441
H4 C8 C11 126.490 H5 C10 C7 125.987
H5 C10 C11 124.877 H6 C11 C8 125.987
H6 C11 C10 124.877 C7 C9 C8 103.591
C7 C10 C11 109.136 C8 C11 C10 109.136
C9 C7 C10 109.069 C9 C8 C11 109.069
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability