return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (Z)-)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-155.612962
Energy at 298.15K-155.620670
Nuclear repulsion energy116.685589
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3189 3023 54.91      
2 A 3172 3007 0.01      
3 A 3081 2921 9.19      
4 A 1712 1623 0.95      
5 A 1599 1516 6.27      
6 A 1501 1423 8.13      
7 A 1353 1282 0.41      
8 A 1071 1015 1.63      
9 A 856 811 0.47      
10 A 311 295 0.03      
11 A 3132 2969 0.00      
12 A 1590 1507 0.00      
13 A 1125 1066 0.00      
14 A 1025 971 0.00      
15 A 387 367 0.00      
16 A 122 116 0.00      
17 A 3132 2970 39.86      
18 A 1594 1511 11.50      
19 A 1134 1075 0.48      
20 A 719 682 45.88      
21 A 100 94 0.68      
22 A 3168 3004 21.54      
23 A 3150 2986 0.00      
24 A 3078 2918 23.65      
25 A 1595 1512 3.89      
26 A 1490 1412 5.74      
27 A 1481 1404 0.00      
28 A 1195 1133 3.30      
29 A 979 928 8.43      
30 A 590 559 6.60      

Unscaled Zero Point Vibrational Energy (zpe) 24313.5 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 23049.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.53057 0.16797 0.13399

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.673 0.677
C2 0.000 -0.673 0.677
C3 0.000 1.595 -0.532
C4 0.000 -1.595 -0.532
H5 0.000 1.184 1.640
H6 0.000 -1.184 1.640
H7 0.000 1.035 -1.471
H8 0.000 -1.035 -1.471
H9 -0.886 2.242 -0.520
H10 0.886 2.242 -0.520
H11 0.886 -2.242 -0.520
H12 -0.886 -2.242 -0.520

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.34561.52102.57031.08972.09132.17802.74372.16332.16333.27333.2733
C21.34562.57031.52102.09131.08972.74372.17803.27333.27332.16332.1633
C31.52102.57033.19072.21033.52711.09322.79251.09681.09683.93863.9386
C42.57031.52103.19073.52712.21032.79251.09323.93863.93861.09681.0968
H51.08972.09132.21033.52712.36793.11373.82032.56242.56244.14554.1455
H62.09131.08973.52712.21032.36793.82033.11374.14554.14552.56242.5624
H72.17802.74371.09322.79253.11373.82032.06941.77401.77403.52533.5253
H82.74372.17802.79251.09323.82033.11372.06943.52533.52531.77401.7740
H92.16333.27331.09683.93862.56244.14551.77403.52531.77114.82194.4848
H102.16333.27331.09683.93862.56244.14551.77403.52531.77114.48484.8219
H113.27332.16333.93861.09684.14552.56243.52531.77404.82194.48481.7711
H123.27332.16333.93861.09684.14552.56243.52531.77404.48484.82191.7711

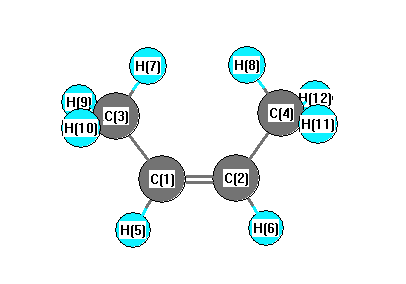
picture of 2-Butene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.342 C1 C2 H6 117.974
C1 C3 H7 111.802 C1 C3 H9 110.408
C1 C3 H10 110.408 C2 C1 C3 127.342
C2 C1 H5 117.974 C2 C4 H8 111.802
C2 C4 H11 110.408 C2 C4 H12 110.408
C3 C1 H5 114.684 C4 C2 H6 114.684
H7 C3 H9 108.203 H7 C3 H10 108.203
H8 C4 H11 108.203 H8 C4 H12 108.203
H9 C3 H10 107.683 H11 C4 H12 107.683
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability