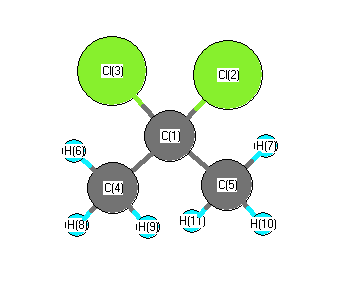
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3166 |
3034 |
6.12 |
65.06 |
0.72 |
0.84 |
| 2 |
A1 |
3068 |
2940 |
11.09 |
302.80 |
0.00 |
0.01 |
| 3 |
A1 |
1487 |
1425 |
4.21 |
7.52 |
0.73 |
0.84 |
| 4 |
A1 |
1420 |
1361 |
7.40 |
0.94 |
0.18 |
0.31 |
| 5 |
A1 |
1184 |
1135 |
35.06 |
3.65 |
0.39 |
0.56 |
| 6 |
A1 |
933 |
894 |
8.88 |
5.50 |
0.67 |
0.80 |
| 7 |
A1 |
569 |
545 |
15.29 |
20.24 |
0.02 |
0.04 |
| 8 |
A1 |
362 |
347 |
0.84 |
2.30 |
0.23 |
0.37 |
| 9 |
A1 |
253 |
243 |
1.17 |
2.36 |
0.64 |
0.78 |
| 10 |
A2 |
3140 |
3009 |
0.00 |
14.22 |
0.75 |
0.86 |
| 11 |
A2 |
1476 |
1414 |
0.00 |
8.61 |
0.75 |
0.86 |
| 12 |
A2 |
1025 |
982 |
0.00 |
0.12 |
0.75 |
0.86 |
| 13 |
A2 |
281 |
269 |
0.00 |
1.14 |
0.75 |
0.86 |
| 14 |
A2 |
265 |
254 |
0.00 |
0.04 |
0.75 |
0.86 |
| 15 |
B1 |
3146 |
3014 |
15.71 |
112.29 |
0.75 |
0.86 |
| 16 |
B1 |
1498 |
1435 |
5.78 |
0.09 |
0.75 |
0.86 |
| 17 |
B1 |
1137 |
1089 |
83.59 |
4.15 |
0.75 |
0.86 |
| 18 |
B1 |
660 |
633 |
92.36 |
9.98 |
0.75 |
0.86 |
| 19 |
B1 |
359 |
344 |
3.53 |
1.26 |
0.75 |
0.86 |
| 20 |
B1 |
296 |
284 |
0.01 |
0.00 |
0.75 |
0.86 |
| 21 |
B2 |
3165 |
3032 |
3.50 |
29.70 |
0.75 |
0.86 |
| 22 |
B2 |
3064 |
2936 |
3.55 |
4.43 |
0.75 |
0.86 |
| 23 |
B2 |
1471 |
1409 |
4.02 |
0.06 |
0.75 |
0.86 |
| 24 |
B2 |
1405 |
1346 |
18.87 |
0.37 |
0.75 |
0.86 |
| 25 |
B2 |
1224 |
1173 |
5.59 |
1.98 |
0.75 |
0.86 |
| 26 |
B2 |
954 |
914 |
0.00 |
0.80 |
0.75 |
0.86 |
| 27 |
B2 |
388 |
372 |
2.63 |
0.85 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 18698.0 cm
-1
Scaled (by 0.9581) Zero Point Vibrational Energy (zpe) 17914.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.924 |
|
|
|
| 2 |
Cl |
-0.487 |
|
|
|
| 3 |
Cl |
-0.487 |
|
|
|
| 4 |
C |
-0.720 |
|
|
|
| 5 |
C |
-0.720 |
|
|
|
| 6 |
H |
0.293 |
|
|
|
| 7 |
H |
0.293 |
|
|
|
| 8 |
H |
0.226 |
|
|
|
| 9 |
H |
0.226 |
|
|
|
| 10 |
H |
0.226 |
|
|
|
| 11 |
H |
0.226 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.363 |
2.363 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.283 |
0.000 |
0.000 |
| y |
0.000 |
-43.198 |
0.000 |
| z |
0.000 |
0.000 |
-43.658 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.855 |
0.000 |
0.000 |
| y |
0.000 |
2.272 |
0.000 |
| z |
0.000 |
0.000 |
1.583 |
|
| Polar |
|---|
| 3z2-r2 | 3.165 |
|---|
| x2-y2 | -4.085 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.871 |
0.000 |
0.000 |
| y |
0.000 |
9.150 |
0.000 |
| z |
0.000 |
0.000 |
9.845 |
<r2> (average value of r
2) Å
2
| <r2> |
186.394 |
| (<r2>)1/2 |
13.653 |