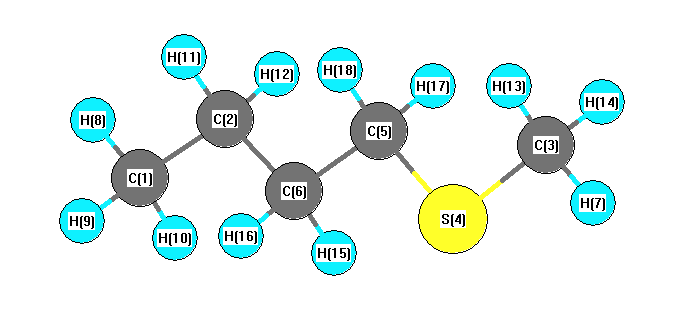
Vibrational Frequencies calculated at mPW1PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3177 |
3045 |
4.75 |
|
|
|
| 2 |
A' |
3145 |
3014 |
29.94 |
|
|
|
| 3 |
A' |
3057 |
2930 |
2.44 |
|
|
|
| 4 |
A' |
3056 |
2929 |
67.98 |
|
|
|
| 5 |
A' |
3054 |
2927 |
58.14 |
|
|
|
| 6 |
A' |
3047 |
2920 |
11.17 |
|
|
|
| 7 |
A' |
3044 |
2917 |
4.23 |
|
|
|
| 8 |
A' |
1489 |
1427 |
5.23 |
|
|
|
| 9 |
A' |
1478 |
1416 |
0.56 |
|
|
|
| 10 |
A' |
1471 |
1409 |
1.02 |
|
|
|
| 11 |
A' |
1465 |
1404 |
0.19 |
|
|
|
| 12 |
A' |
1457 |
1396 |
11.81 |
|
|
|
| 13 |
A' |
1407 |
1348 |
3.11 |
|
|
|
| 14 |
A' |
1394 |
1336 |
3.16 |
|
|
|
| 15 |
A' |
1338 |
1282 |
3.06 |
|
|
|
| 16 |
A' |
1321 |
1266 |
5.16 |
|
|
|
| 17 |
A' |
1233 |
1181 |
18.94 |
|
|
|
| 18 |
A' |
1132 |
1085 |
1.46 |
|
|
|
| 19 |
A' |
1094 |
1049 |
0.76 |
|
|
|
| 20 |
A' |
1056 |
1012 |
0.16 |
|
|
|
| 21 |
A' |
973 |
932 |
6.09 |
|
|
|
| 22 |
A' |
911 |
873 |
0.85 |
|
|
|
| 23 |
A' |
785 |
752 |
2.10 |
|
|
|
| 24 |
A' |
737 |
706 |
2.65 |
|
|
|
| 25 |
A' |
422 |
404 |
0.34 |
|
|
|
| 26 |
A' |
316 |
303 |
0.46 |
|
|
|
| 27 |
A' |
237 |
227 |
0.01 |
|
|
|
| 28 |
A' |
116 |
112 |
0.83 |
|
|
|
| 29 |
A" |
3152 |
3020 |
9.50 |
|
|
|
| 30 |
A" |
3139 |
3008 |
46.53 |
|
|
|
| 31 |
A" |
3114 |
2984 |
44.09 |
|
|
|
| 32 |
A" |
3094 |
2965 |
0.77 |
|
|
|
| 33 |
A" |
3078 |
2950 |
3.41 |
|
|
|
| 34 |
A" |
1476 |
1414 |
7.24 |
|
|
|
| 35 |
A" |
1444 |
1384 |
8.27 |
|
|
|
| 36 |
A" |
1325 |
1269 |
0.26 |
|
|
|
| 37 |
A" |
1298 |
1244 |
0.18 |
|
|
|
| 38 |
A" |
1220 |
1169 |
0.30 |
|
|
|
| 39 |
A" |
1066 |
1022 |
0.39 |
|
|
|
| 40 |
A" |
964 |
924 |
4.31 |
|
|
|
| 41 |
A" |
919 |
881 |
0.53 |
|
|
|
| 42 |
A" |
794 |
761 |
0.00 |
|
|
|
| 43 |
A" |
747 |
716 |
4.80 |
|
|
|
| 44 |
A" |
254 |
244 |
0.04 |
|
|
|
| 45 |
A" |
180 |
173 |
0.64 |
|
|
|
| 46 |
A" |
132 |
126 |
0.01 |
|
|
|
| 47 |
A" |
93 |
90 |
0.61 |
|
|
|
| 48 |
A" |
57 |
55 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35479.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33999.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.071 |
|
|
|
| 2 |
C |
-0.109 |
|
|
|
| 3 |
C |
-0.179 |
|
|
|
| 4 |
S |
-0.071 |
|
|
|
| 5 |
C |
-0.157 |
|
|
|
| 6 |
C |
-0.065 |
|
|
|
| 7 |
H |
0.078 |
|
|
|
| 8 |
H |
0.040 |
|
|
|
| 9 |
H |
0.043 |
|
|
|
| 10 |
H |
0.043 |
|
|
|
| 11 |
H |
0.036 |
|
|
|
| 12 |
H |
0.036 |
|
|
|
| 13 |
H |
0.074 |
|
|
|
| 14 |
H |
0.074 |
|
|
|
| 15 |
H |
0.048 |
|
|
|
| 16 |
H |
0.048 |
|
|
|
| 17 |
H |
0.066 |
|
|
|
| 18 |
H |
0.066 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.075 |
1.133 |
0.000 |
1.562 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.316 |
3.000 |
0.000 |
| y |
3.000 |
-44.398 |
0.000 |
| z |
0.000 |
0.000 |
-47.654 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.290 |
3.000 |
0.000 |
| y |
3.000 |
3.087 |
0.000 |
| z |
0.000 |
0.000 |
-1.797 |
|
| Polar |
|---|
| 3z2-r2 | -3.594 |
|---|
| x2-y2 | -2.918 |
|---|
| xy | 3.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
370.361 |
| (<r2>)1/2 |
19.245 |