return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (Thiophene, tetrahydro-3-methyl-)

using model chemistry: MP2/CEP-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/CEP-31G*
 hartrees
Energy at 0K-43.942945
Energy at 298.15K-43.954865
HF Energy-43.200247
Nuclear repulsion energy136.507449
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3191 3030 25.59      
2 A 3183 3022 33.10      
3 A 3181 3020 20.05      
4 A 3173 3013 29.36      
5 A 3162 3002 15.13      
6 A 3116 2958 25.22      
7 A 3111 2953 71.20      
8 A 3099 2943 20.51      
9 A 3089 2932 16.64      
10 A 3079 2923 27.29      
11 A 1540 1462 3.58      
12 A 1536 1458 3.46      
13 A 1520 1443 5.76      
14 A 1514 1437 6.72      
15 A 1509 1432 2.72      
16 A 1447 1374 9.31      
17 A 1393 1322 4.82      
18 A 1371 1302 0.61      
19 A 1342 1274 5.16      
20 A 1323 1256 4.36      
21 A 1297 1231 22.09      
22 A 1261 1198 3.67      
23 A 1207 1146 2.24      
24 A 1152 1093 0.12      
25 A 1099 1043 1.59      
26 A 1091 1036 0.26      
27 A 1059 1006 0.55      
28 A 1021 970 2.65      
29 A 987 937 0.38      
30 A 956 908 0.76      
31 A 926 879 4.71      
32 A 882 837 1.69      
33 A 821 779 2.00      
34 A 742 705 4.97      
35 A 717 681 2.52      
36 A 620 588 1.09      
37 A 498 473 0.11      
38 A 384 364 0.09      
39 A 352 334 0.14      
40 A 243 230 0.11      
41 A 188 179 0.16      
42 A 119 113 2.18      

Unscaled Zero Point Vibrational Energy (zpe) 31748.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 30141.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/CEP-31G*
ABC
0.13960 0.08563 0.07037

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/CEP-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
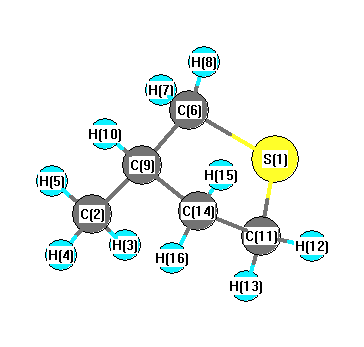
S1 1.477 0.945 -0.538
C2 -1.733 -0.171 -0.717
H3 -1.130 -0.109 -1.639
H4 -2.236 -1.156 -0.691
H5 -2.514 0.610 -0.771
C6 -0.030 1.333 0.465
H7 -0.587 2.151 -0.024
H8 0.279 1.658 1.474
C9 -0.860 0.029 0.542
H10 -1.517 0.076 1.432
C11 1.262 -0.884 -0.375
H12 2.226 -1.337 -0.087
H13 0.941 -1.308 -1.341
C14 0.185 -1.094 0.717
H15 0.652 -1.010 1.716
H16 -0.275 -2.097 0.629

Atom - Atom Distances (Å)
  S1 C2 H3 H4 H5 C6 H7 H8 C9 H10 C11 H12 H13 C14 H15 H16
S13.40363.01974.26874.01181.85122.44502.44802.73243.68801.84862.44332.45132.72083.09583.6988
C23.40361.10381.10581.10552.56172.68083.49221.54472.17423.09784.17492.97262.56713.50882.7652
H33.01971.10381.79381.78482.77802.83043.84692.20133.10112.81383.89532.41152.87243.90393.1344
H44.26871.10581.79381.78933.52113.75494.35072.19452.55783.52244.50583.24662.80143.76192.5442
H54.01181.10551.78481.78932.86772.57833.73372.18982.47714.07985.16933.99283.52224.33983.7816
C61.85122.56172.77803.52112.86771.10381.10411.54772.17442.69973.53843.34392.45002.74243.4423
H72.44502.68082.83043.75492.57831.10381.79912.21262.70043.57084.48094.00453.41733.81514.3087
H82.44803.49223.84694.35073.73371.10411.79912.19542.39363.29353.89844.14302.85612.70503.8881
C92.73241.54472.20132.19452.18981.54772.21262.19541.10782.48543.43292.92901.54492.17862.2068
H103.68802.17423.10112.55782.47712.17442.70042.39361.10783.45104.27933.95622.18602.44202.6283
C111.84863.09782.81383.52244.07982.69973.57083.29352.48543.45101.10311.10301.54802.18182.2000
H122.44334.17493.89534.50585.16933.53844.48093.89843.43294.27931.10311.79522.20672.41582.7098
H132.45132.97262.41153.24663.99283.34394.00454.14302.92903.95621.10301.79522.20333.08532.4459
C142.72082.56712.87242.80143.52222.45003.41732.85611.54492.18601.54802.20672.20331.10561.1062
H153.09583.50883.90393.76194.33982.74243.81512.70502.17862.44202.18182.41583.08531.10561.7943
H163.69882.76523.13442.54423.78163.44234.30873.88812.20682.62832.20002.70982.44591.10621.7943

picture of Thiophene, tetrahydro-3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C6 H7 109.033 S1 C6 H8 109.239
S1 C6 C9 106.672 S1 C11 H12 109.113
S1 C11 H13 109.704 S1 C11 C14 106.123
C2 C9 C6 111.871 C2 C9 H10 108.995
C2 C9 C14 112.380 H3 C2 H4 108.547
H3 C2 H5 107.774 H3 C2 C9 111.348
H4 C2 H5 108.030 H4 C2 C9 110.688
H5 C2 C9 110.339 C6 S1 C11 93.723
C6 C9 H10 108.812 C6 C9 C14 104.786
H7 C6 H8 109.143 H7 C6 C9 112.033
H8 C6 C9 110.651 C9 C14 C11 106.946
C9 C14 H15 109.449 C9 C14 H16 111.626
H10 C9 C14 109.890 C11 C14 H15 109.485
C11 C14 H16 110.870 H12 C11 H13 108.931
H12 C11 C14 111.589 H13 C11 C14 111.324
H15 C14 H16 108.441
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability