All results from a given calculation for CH3CO (Acetyl radical)
using model chemistry: MP2/SDD
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
2A' |
Energy calculated at MP2/SDD
| | hartrees |
|---|
| Energy at 0K | -152.531525 |
| Energy at 298.15K | |
| HF Energy | -152.246118 |
| Nuclear repulsion energy | 61.524954 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/SDD
Geometric Data calculated at MP2/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.993 |
-0.682 |
0.000 |
| C2 |
0.000 |
0.513 |
0.000 |
| O3 |
1.212 |
0.479 |
0.000 |
| H4 |
-0.474 |
-1.655 |
0.000 |
| H5 |
-1.633 |
-0.581 |
0.891 |
| H6 |
-1.633 |
-0.581 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
| C1 | | 1.5537 | 2.4924 | 1.1030 | 1.1017 | 1.1017 |
C2 | 1.5537 | | 1.2127 | 2.2193 | 2.1581 | 2.1581 | O3 | 2.4924 | 1.2127 | | 2.7202 | 3.1646 | 3.1646 | H4 | 1.1030 | 2.2193 | 2.7202 | | 1.8145 | 1.8145 | H5 | 1.1017 | 2.1581 | 3.1646 | 1.8145 | | 1.7817 | H6 | 1.1017 | 2.1581 | 3.1646 | 1.8145 | 1.7817 | |
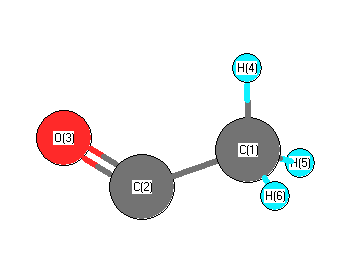 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
128.146 |
|
C2 |
C1 |
H4 |
112.193 |
| C2 |
C1 |
H5 |
107.500 |
|
C2 |
C1 |
H6 |
107.500 |
| H4 |
C1 |
H5 |
110.775 |
|
H4 |
C1 |
H6 |
110.775 |
| H5 |
C1 |
H6 |
107.914 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
