return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CClCHCH2 (1,3-Butadiene, 2-chloro-)

using model chemistry: MP2/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/aug-cc-pVDZ
 hartrees
Energy at 0K-614.562956
Energy at 298.15K 
HF Energy-613.860290
Nuclear repulsion energy196.120359
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3306 3171 1.05 55.03 0.75 0.85
2 A' 3301 3165 2.69 62.83 0.65 0.79
3 A' 3201 3069 4.73 198.03 0.12 0.21
4 A' 3192 3061 0.81 60.73 0.13 0.23
5 A' 3188 3057 1.20 54.96 0.20 0.33
6 A' 1683 1614 4.83 170.64 0.20 0.33
7 A' 1620 1554 33.15 3.63 0.32 0.48
8 A' 1450 1390 1.05 46.79 0.39 0.56
9 A' 1399 1342 4.70 3.07 0.49 0.66
10 A' 1303 1250 0.12 25.92 0.22 0.36
11 A' 1243 1192 43.85 8.06 0.53 0.70
12 A' 1025 983 3.19 3.52 0.56 0.72
13 A' 912 874 7.42 0.47 0.69 0.81
14 A' 645 618 19.25 12.64 0.06 0.11
15 A' 521 500 0.97 5.25 0.45 0.62
16 A' 386 370 1.62 2.44 0.75 0.86
17 A' 248 238 0.06 2.06 0.65 0.79
18 A" 992 952 21.59 0.17 0.75 0.86
19 A" 938 900 32.84 3.70 0.75 0.86
20 A" 891 855 38.77 4.33 0.75 0.86
21 A" 750 719 1.36 0.32 0.75 0.86
22 A" 663 636 0.05 0.05 0.75 0.86
23 A" 411 394 9.77 0.22 0.75 0.86
24 A" 154 147 0.39 0.23 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 16711.1 cm-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 16025.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/aug-cc-pVDZ
ABC
0.17798 0.12311 0.07277

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.316 1.902 0.000
C2 0.000 0.584 0.000
C3 1.378 0.081 0.000
C4 1.747 -1.222 0.000
Cl5 -1.291 -0.606 0.000
H6 0.491 2.639 0.000
H7 -1.351 2.246 0.000
H8 2.144 0.865 0.000
H9 2.805 -1.494 0.000
H10 1.006 -2.024 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 Cl5 H6 H7 H8 H9 H10
C11.35472.48673.74352.69121.09271.09072.66944.61204.1422
C21.35471.46702.51311.75652.11272.14132.16253.49112.7955
C32.48671.46701.35432.75652.70773.48311.09632.12552.1374
C43.74352.51311.35433.10064.06054.65012.12441.09201.0919
Cl52.69121.75652.75653.10063.70262.85293.73744.19152.6998
H61.09272.11272.70774.06053.70261.88272.42524.73724.6914
H71.09072.14133.48314.65012.85291.88273.75775.59084.8771
H82.66942.16251.09632.12443.73742.42523.75772.45013.1049
H94.61203.49112.12551.09204.19154.73725.59082.45011.8749
H104.14222.79552.13741.09192.69984.69144.87713.10491.8749

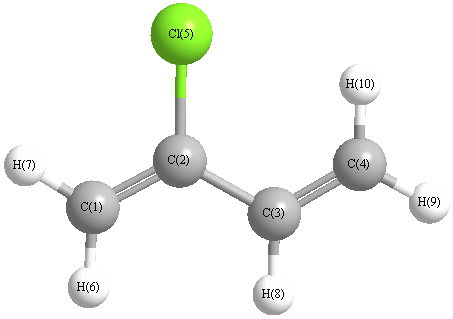
picture of 1,3-Butadiene, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.539 C1 C2 Cl5 119.198
C2 C1 H6 118.972 C2 C1 H7 121.872
C2 C3 C4 125.893 C2 C3 H8 114.275
C3 C2 Cl5 117.263 C3 C4 H9 120.270
C3 C4 H10 121.428 C4 C3 H8 119.832
H6 C1 H7 119.157 H9 C4 H10 118.302
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability