return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3OCH2 (methoxymethyl radical)

using model chemistry: MP2/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A
Energy calculated at MP2/aug-cc-pVDZ
 hartrees
Energy at 0K-153.933806
Energy at 298.15K-153.938793
HF Energy-153.459479
Nuclear repulsion energy76.843035
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3321 3185 9.55      
2 A 3203 3072 15.16      
3 A 3154 3025 24.39      
4 A 3139 3011 36.70      
5 A 3052 2927 50.09      
6 A 1498 1436 16.64      
7 A 1490 1429 4.75      
8 A 1483 1422 5.24      
9 A 1448 1388 1.17      
10 A 1270 1218 144.04      
11 A 1247 1196 60.14      
12 A 1167 1119 2.27      
13 A 1131 1085 8.15      
14 A 959 920 31.94      
15 A 656 629 38.15      
16 A 435 418 3.49      
17 A 303 290 7.42      
18 A 175 168 1.70      

Unscaled Zero Point Vibrational Energy (zpe) 14565.8 cm-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 13968.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/aug-cc-pVDZ
ABC
1.51545 0.35647 0.30736

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/aug-cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.093 -0.559 -0.043
C2 -1.138 0.175 0.015
C3 1.199 0.236 0.074
H4 -1.945 -0.555 -0.124
H5 -1.244 0.671 0.994
H6 -1.173 0.929 -0.790
H7 2.135 -0.311 -0.024
H8 1.115 1.277 -0.252

Atom - Atom Distances (Å)
  O1 C2 C3 H4 H5 H6 H7 H8
O11.43461.36692.03942.09162.09212.05732.1125
C21.43462.33831.09651.10211.10343.30932.5226
C31.36692.33833.24742.64582.61801.08881.0948
H42.03941.09653.24741.80071.79974.08843.5687
H52.09161.10212.64581.80071.80323.66312.7357
H62.09211.10342.61801.79971.80323.61522.3766
H72.05733.30931.08884.08843.66313.61521.9014
H82.11252.52261.09483.56872.73572.37661.9014

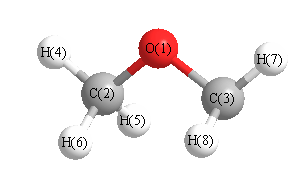
picture of methoxymethyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 H4 106.597 O1 C2 H5 110.401
O1 C2 H6 110.356 O1 C3 H7 113.317
O1 C3 H8 117.789 C2 O1 C3 113.135
H4 C2 H5 109.974 H4 C2 H6 109.776
H5 C2 H6 109.692 H7 C3 H8 121.091
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability