return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-methylbutyraldehyde)

using model chemistry: MP2/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/aug-cc-pVDZ
 hartrees
Energy at 0K-270.983875
Energy at 298.15K-270.994629
HF Energy-270.051403
Nuclear repulsion energy239.109707
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3171 3041 15.95      
2 A 3162 3032 26.42      
3 A 3159 3029 20.03      
4 A 3150 3021 29.14      
5 A 3100 2973 18.96      
6 A 3070 2944 15.54      
7 A 3062 2936 22.58      
8 A 3047 2922 25.69      
9 A 3023 2899 17.08      
10 A 2934 2814 105.56      
11 A 1729 1658 132.32      
12 A 1501 1440 3.03      
13 A 1491 1430 10.18      
14 A 1486 1425 5.92      
15 A 1482 1422 6.60      
16 A 1471 1411 0.12      
17 A 1416 1358 5.91      
18 A 1396 1339 2.06      
19 A 1389 1332 4.47      
20 A 1362 1306 4.05      
21 A 1348 1293 1.80      
22 A 1301 1248 2.08      
23 A 1258 1206 0.90      
24 A 1190 1141 2.30      
25 A 1169 1121 1.99      
26 A 1113 1068 1.91      
27 A 1055 1012 0.48      
28 A 1019 978 11.96      
29 A 967 927 5.65      
30 A 936 898 7.91      
31 A 919 881 11.08      
32 A 797 765 4.34      
33 A 776 744 1.86      
34 A 659 632 7.63      
35 A 406 389 0.73      
36 A 382 366 0.55      
37 A 295 283 3.50      
38 A 263 253 0.70      
39 A 237 227 3.46      
40 A 204 195 0.33      
41 A 95 91 1.76      
42 A 73 70 5.20      

Unscaled Zero Point Vibrational Energy (zpe) 31031.3 cm-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 29759.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/aug-cc-pVDZ
ABC
0.23011 0.07019 0.05872

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/aug-cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.379 -0.684 0.247
C2 -0.083 0.082 0.428
C3 1.034 -0.704 -0.295
C4 2.437 -0.152 -0.016
C5 -0.207 1.532 -0.040
O6 -2.384 -0.246 -0.303
H7 -1.368 -1.731 0.640
H8 0.139 0.045 1.515
H9 0.992 -1.762 0.019
H10 0.829 -0.685 -1.381
H11 3.206 -0.779 -0.494
H12 2.554 0.871 -0.404
H13 2.639 -0.132 1.068
H14 -0.364 1.569 -1.129
H15 -1.067 2.024 0.437
H16 0.700 2.103 0.206

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.51622.47353.86232.52291.22631.11882.10812.61512.74324.64544.27934.13812.82882.73293.4773
C21.51621.54562.56991.52862.43632.23191.10962.17352.16643.52262.87602.80442.17232.17782.1791
C32.47351.54561.53352.56953.44862.77472.15391.10481.10512.18212.19162.18162.79603.52062.8708
C43.86232.56991.53353.13444.83064.17182.76882.16422.17561.10111.10031.10263.47084.14992.8549
C52.52291.52862.56953.13442.82273.52932.17843.50582.79024.14622.86193.47771.10151.10011.0998
O61.22632.43633.44864.83062.82272.03193.12313.71523.41735.61845.06375.20792.83882.72663.9099
H71.11882.23192.77474.17183.52932.03192.48822.44063.16294.80734.82114.33513.87723.77324.3778
H82.10811.10962.15392.76882.17843.12312.48822.49683.06523.75783.19352.54623.09312.55622.5023
H92.61512.17351.10482.16423.50583.71522.44062.49681.77372.47583.09102.54323.77594.33083.8808
H102.74322.16641.10512.17562.79023.41733.16293.06521.77372.53872.52033.09462.56303.77373.2108
H114.64543.52262.18211.10114.14625.61844.80733.75782.47582.53871.77661.78244.31985.19463.8825
H124.27932.87602.19161.10032.86195.06374.82113.19353.09102.52031.77661.78353.08643.89222.3079
H134.13812.80442.18161.10263.47775.20794.33512.54622.54323.09461.78241.78354.09144.33443.0819
H142.82882.17232.79603.47081.10152.83883.87723.09313.77592.56304.31983.08644.09141.77601.7890
H152.73292.17783.52064.14991.10012.72663.77322.55624.33083.77375.19463.89224.33441.77601.7840
H163.47732.17912.87082.85491.09983.90994.37782.50233.88083.21083.88252.30793.08191.78901.7840

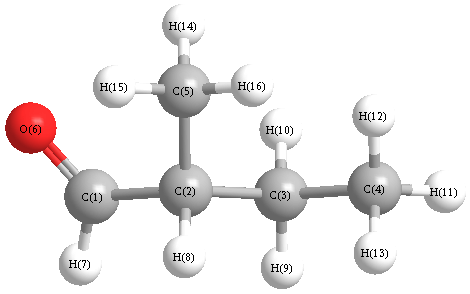
picture of 2-methylbutyraldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 107.773 C1 C2 C5 111.906
C1 C2 H8 105.768 C2 C1 O6 125.005
C2 C1 H7 114.950 C2 C3 C4 113.157
C2 C3 H9 109.055 C2 C3 H10 108.493
C2 C5 H14 110.306 C2 C5 H15 110.832
C2 C5 H16 110.950 C3 C2 C5 113.402
C3 C2 H8 107.290 C3 C4 H11 110.771
C3 C4 H12 111.576 C3 C4 H13 110.642
C4 C3 H9 109.145 C4 C3 H10 110.025
C5 C2 H8 110.310 O6 C1 H7 120.033
H9 C3 H10 106.763 H11 C4 H12 107.620
H11 C4 H13 107.967 H12 C4 H13 108.123
H14 C5 H15 107.553 H14 C5 H16 108.721
H15 C5 H16 108.378
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability