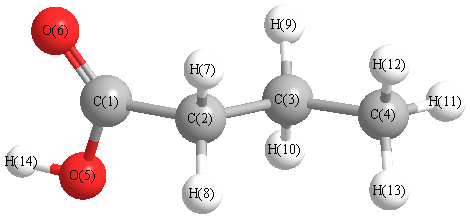return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Butanoic acid)

using model chemistry: MP2/aug-cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/aug-cc-pVTZ
 hartrees
Energy at 0K-307.171933
Energy at 298.15K-307.181359
HF Energy-306.000129
Nuclear repulsion energy239.538623
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/aug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3747 3570 77.37      
2 A 3171 3021 15.04      
3 A 3160 3011 21.52      
4 A 3154 3006 19.35      
5 A 3125 2978 0.74      
6 A 3101 2955 12.85      
7 A 3078 2933 10.82      
8 A 3067 2922 19.01      
9 A 1803 1718 291.14      
10 A 1525 1453 8.04      
11 A 1519 1448 8.19      
12 A 1512 1441 1.85      
13 A 1495 1425 7.18      
14 A 1419 1353 2.47      
15 A 1390 1324 50.41      
16 A 1371 1307 31.35      
17 A 1334 1271 6.38      
18 A 1293 1233 1.58      
19 A 1265 1206 7.24      
20 A 1204 1147 136.30      
21 A 1125 1072 7.17      
22 A 1086 1034 93.63      
23 A 1073 1022 18.89      
24 A 934 890 4.17      
25 A 900 857 0.68      
26 A 883 842 12.92      
27 A 760 724 4.78      
28 A 744 709 26.56      
29 A 620 590 72.49      
30 A 583 556 50.93      
31 A 432 412 2.75      
32 A 331 315 0.99      
33 A 252 240 0.02      
34 A 183 174 0.07      
35 A 93 89 0.36      
36 A 36 34 0.68      

Unscaled Zero Point Vibrational Energy (zpe) 26383.0 cm-1
Scaled (by 0.9529) Zero Point Vibrational Energy (zpe) 25140.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/aug-cc-pVTZ
ABC
0.26963 0.06280 0.05637

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/aug-cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.163 -0.134 0.111
C2 -0.242 -0.191 0.631
C3 -1.241 0.088 -0.498
C4 -2.677 0.008 0.003
O5 1.604 1.147 0.010
O6 1.836 -1.081 -0.229
H7 -0.407 -1.187 1.039
H8 -0.368 0.548 1.422
H9 -1.082 -0.638 -1.297
H10 -1.042 1.076 -0.914
H11 -3.385 0.203 -0.800
H12 -2.891 -0.981 0.409
H13 -2.852 0.740 0.792
H14 2.492 1.090 -0.376

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13 H14
C11.49902.48933.84321.35911.21122.10562.12722.69772.71574.65054.15224.16521.8718
C21.49901.53262.52172.36372.41901.08841.08992.15022.15223.47592.77302.77613.1834
C32.48931.53261.52253.07803.30262.16352.15821.09161.09082.16862.16512.16483.8673
C43.84322.52171.52254.42954.64812.76592.76282.15672.15731.08921.09021.09035.2944
O51.35912.36373.07804.42952.25353.24852.49823.48022.80335.14194.98964.54290.9702
O61.21122.41903.30264.64812.25352.57923.19973.13913.66145.40724.77155.13262.2730
H72.10561.08842.16352.76593.24852.57921.77702.49313.05573.76612.57113.12303.9488
H82.12721.08992.15822.76282.49823.19971.77703.05142.48753.76313.11932.57033.4214
H92.69772.15021.09162.15673.48023.13912.49313.05141.75702.50162.51063.06584.0757
H102.71572.15221.09082.15732.80333.66143.05572.48751.75702.50323.06622.51033.5748
H114.65053.47592.16861.08925.14195.40723.76613.76312.50162.50321.76301.76315.9591
H124.15222.77302.16511.09024.98964.77152.57113.11932.51063.06621.76301.76325.8213
H134.16522.77612.16481.09034.54295.13263.12302.57033.06582.51031.76311.76325.4821
H141.87183.18343.86735.29440.97022.27303.94883.42144.07573.57485.95915.82135.4821

picture of Butanoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 110.392 C1 C2 H7 107.879
C1 C2 H8 109.483 C1 O5 H14 105.734
C2 C1 O5 111.488 C2 C1 O6 126.066
C2 C3 C4 111.259 C2 C3 H9 108.889
C2 C3 H10 109.090 C3 C2 H7 110.116
C3 C2 H8 109.612 C3 C4 H11 111.181
C3 C4 H12 110.834 C3 C4 H13 110.806
C4 C3 H9 110.091 C4 C3 H10 110.177
O5 C1 O6 122.402 H7 C2 H8 109.330
H9 C3 H10 107.237 H11 C4 H12 107.981
H11 C4 H13 107.989 H12 C4 H13 107.919
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability