return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2C(CH3)OCH3 (1-Propene, 2-methoxy-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-231.838091
Energy at 298.15K-231.846704
HF Energy-230.978619
Nuclear repulsion energy180.577688
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3345 3159 2.12      
2 A 3237 3058 0.09      
3 A 3218 3040 4.78      
4 A 3215 3036 19.51      
5 A 3182 3006 7.45      
6 A 3164 2988 30.77      
7 A 3103 2931 11.26      
8 A 3080 2909 40.30      
9 A 1732 1636 80.97      
10 A 1542 1456 8.34      
11 A 1533 1448 7.95      
12 A 1517 1433 10.02      
13 A 1506 1422 5.13      
14 A 1503 1420 3.76      
15 A 1443 1363 6.34      
16 A 1418 1339 33.74      
17 A 1328 1254 152.68      
18 A 1225 1157 0.59      
19 A 1194 1127 3.11      
20 A 1146 1083 89.01      
21 A 1072 1013 1.23      
22 A 1029 971 3.78      
23 A 948 895 5.08      
24 A 864 816 48.17      
25 A 828 782 7.94      
26 A 763 721 2.72      
27 A 540 510 4.87      
28 A 476 449 3.79      
29 A 417 394 0.74      
30 A 323 305 0.99      
31 A 253 239 0.92      
32 A 210 198 1.08      
33 A 84 79 6.56      

Unscaled Zero Point Vibrational Energy (zpe) 25216.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 23817.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.27311 0.14453 0.10138

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.804 -0.081 0.197
H2 1.744 -0.005 1.287
H3 2.638 -0.729 -0.067
H4 1.984 0.912 -0.222
O5 0.647 -0.693 -0.347
C6 -0.582 1.413 -0.043
H7 0.002 1.836 0.778
H8 -0.176 1.812 -0.976
H9 -1.610 1.755 0.064
C10 -0.545 -0.086 -0.048
C11 -1.617 -0.860 0.143
H12 -2.591 -0.418 0.291
H13 -1.525 -1.936 0.124

Atom - Atom Distances (Å)
  C1 H2 H3 H4 O5 C6 H7 H8 H9 C10 C11 H12 H13
C11.09441.08851.09301.41702.82532.69452.97973.87892.36203.50954.40913.8118
H21.09441.77681.78212.08403.03072.58473.47913.97982.65083.65174.46663.9704
H31.08851.77681.77342.01073.86753.77383.89884.92293.24774.26295.25074.3389
H41.09301.78211.77342.09302.62072.40492.45853.70282.72514.03074.79244.5330
O51.41702.08402.01072.09302.45772.84212.71093.35571.37112.32323.31212.5467
C62.82533.03073.86752.62072.45771.09191.09281.08881.49992.50462.73893.4834
H72.69452.58473.77382.40492.84211.09191.76281.76432.16233.20793.46974.1212
H82.97973.47913.89882.45852.71091.09281.76281.77212.14453.23513.52254.1321
H93.87893.97984.92293.70283.35571.08881.76431.77212.13032.61652.39553.6930
C102.36202.65083.24772.72511.37111.49992.16232.14452.13031.33592.10022.1003
C113.50953.65174.26294.03072.32322.50463.20793.23512.61651.33591.07931.0804
H124.40914.46665.25074.79243.31212.73893.46973.52252.39552.10021.07931.8625
H133.81183.97044.33894.53302.54673.48344.12124.13213.69302.10031.08041.8625

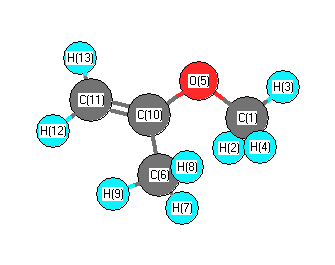
picture of 1-Propene, 2-methoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O5 C10 115.801 H2 C1 H3 108.965
H2 C1 H4 109.122 H2 C1 O5 111.515
H3 C1 H4 108.771 H3 C1 O5 105.992
H4 C1 O5 112.355 O5 C10 C6 117.685
O5 C10 C11 118.228 C6 C10 C11 123.968
H7 C6 H8 107.585 H7 C6 H9 108.008
H7 C6 C10 112.124 H8 C6 H9 108.641
H8 C6 C10 110.635 H9 C6 C10 109.739
C10 C11 H12 120.442 C10 C11 H13 120.372
H12 C11 H13 119.165
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability