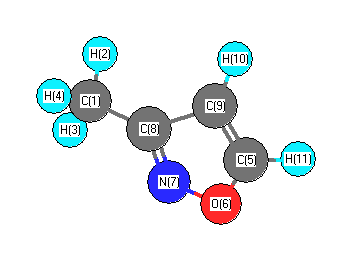
Vibrational Frequencies calculated at MP2/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3323 |
3172 |
0.33 |
|
|
|
| 2 |
A |
3297 |
3147 |
0.09 |
|
|
|
| 3 |
A |
3177 |
3032 |
4.40 |
|
|
|
| 4 |
A |
3162 |
3018 |
10.31 |
|
|
|
| 5 |
A |
3094 |
2953 |
9.82 |
|
|
|
| 6 |
A |
1598 |
1525 |
9.16 |
|
|
|
| 7 |
A |
1589 |
1517 |
8.87 |
|
|
|
| 8 |
A |
1534 |
1464 |
12.96 |
|
|
|
| 9 |
A |
1496 |
1428 |
5.13 |
|
|
|
| 10 |
A |
1427 |
1362 |
18.80 |
|
|
|
| 11 |
A |
1352 |
1290 |
12.67 |
|
|
|
| 12 |
A |
1295 |
1237 |
1.64 |
|
|
|
| 13 |
A |
1154 |
1102 |
9.97 |
|
|
|
| 14 |
A |
1132 |
1081 |
2.88 |
|
|
|
| 15 |
A |
1050 |
1002 |
11.84 |
|
|
|
| 16 |
A |
1014 |
968 |
5.18 |
|
|
|
| 17 |
A |
973 |
928 |
1.88 |
|
|
|
| 18 |
A |
917 |
876 |
0.35 |
|
|
|
| 19 |
A |
857 |
818 |
0.33 |
|
|
|
| 20 |
A |
850 |
811 |
9.71 |
|
|
|
| 21 |
A |
775 |
740 |
57.37 |
|
|
|
| 22 |
A |
657 |
627 |
0.28 |
|
|
|
| 23 |
A |
637 |
608 |
5.67 |
|
|
|
| 24 |
A |
584 |
558 |
5.14 |
|
|
|
| 25 |
A |
336 |
321 |
5.18 |
|
|
|
| 26 |
A |
264 |
252 |
4.91 |
|
|
|
| 27 |
A |
134 |
128 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18838.5 cm
-1
Scaled (by 0.9545) Zero Point Vibrational Energy (zpe) 17981.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.579 |
|
|
-0.560 |
| 2 |
H |
0.211 |
|
|
0.156 |
| 3 |
H |
0.225 |
|
|
0.157 |
| 4 |
H |
0.211 |
|
|
0.156 |
| 5 |
C |
0.118 |
|
|
0.218 |
| 6 |
O |
-0.323 |
|
|
-0.151 |
| 7 |
N |
-0.201 |
|
|
-0.346 |
| 8 |
C |
0.217 |
|
|
0.617 |
| 9 |
C |
-0.338 |
|
|
-0.599 |
| 10 |
H |
0.215 |
|
|
0.243 |
| 11 |
H |
0.244 |
|
|
0.110 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.771 |
3.017 |
0.000 |
3.114 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.742 |
2.979 |
0.000 |
3.070 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.713 |
2.465 |
-0.000 |
| y |
2.465 |
-36.928 |
0.000 |
| z |
-0.000 |
0.000 |
-37.118 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.310 |
2.465 |
-0.000 |
| y |
2.465 |
-3.012 |
0.000 |
| z |
-0.000 |
0.000 |
-3.298 |
|
| Polar |
|---|
| 3z2-r2 | -6.595 |
|---|
| x2-y2 | 6.215 |
|---|
| xy | 2.465 |
|---|
| xz | -0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.494 |
0.147 |
0.000 |
| y |
0.147 |
6.615 |
0.000 |
| z |
0.000 |
0.000 |
3.116 |
<r2> (average value of r
2) Å
2
| <r2> |
142.068 |
| (<r2>)1/2 |
11.919 |