return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Butanenitrile, 2-methyl-)

using model chemistry: MP2/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-311G**
 hartrees
Energy at 0K-249.993923
Energy at 298.15K-250.003416
HF Energy-249.086619
Nuclear repulsion energy219.889272
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3186 3027 14.37      
2 A 3175 3017 18.19      
3 A 3173 3015 27.12      
4 A 3163 3006 25.52      
5 A 3134 2978 7.31      
6 A 3086 2932 12.81      
7 A 3083 2929 21.93      
8 A 3077 2924 2.32      
9 A 3076 2923 12.78      
10 A 2191 2081 0.27      
11 A 1534 1457 1.99      
12 A 1523 1448 11.02      
13 A 1519 1443 9.24      
14 A 1515 1440 6.32      
15 A 1502 1428 0.26      
16 A 1431 1360 2.91      
17 A 1428 1357 5.65      
18 A 1394 1324 0.35      
19 A 1364 1296 0.98      
20 A 1329 1263 1.76      
21 A 1298 1233 1.39      
22 A 1204 1144 1.72      
23 A 1167 1109 0.17      
24 A 1130 1074 3.57      
25 A 1074 1020 1.22      
26 A 1017 966 1.14      
27 A 985 936 3.50      
28 A 919 873 0.26      
29 A 823 782 0.52      
30 A 779 741 2.29      
31 A 558 530 0.07      
32 A 531 505 0.69      
33 A 405 385 0.20      
34 A 318 302 0.53      
35 A 297 282 0.11      
36 A 219 208 0.06      
37 A 201 191 2.01      
38 A 159 151 4.92      
39 A 92 87 1.36      

Unscaled Zero Point Vibrational Energy (zpe) 29027.6 cm-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 27582.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-311G**
ABC
0.21887 0.07341 0.05890

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-311G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
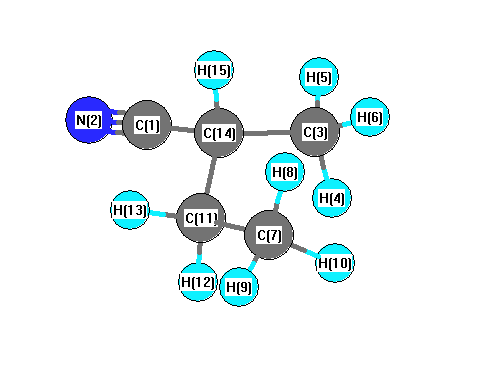
C1 -1.524 -0.279 0.122
N2 -2.611 -0.663 -0.108
C3 -0.003 1.633 -0.152
H4 -0.087 1.643 -1.243
H5 -0.779 2.282 0.260
H6 0.971 2.038 0.129
C7 2.305 -0.419 0.057
H8 2.461 -0.309 1.135
H9 2.977 -1.202 -0.303
H10 2.596 0.517 -0.428
C11 0.852 -0.787 -0.245
H12 0.685 -0.820 -1.328
H13 0.637 -1.789 0.141
C14 -0.158 0.198 0.374
H15 -0.016 0.202 1.462

Atom - Atom Distances (Å)
  C1 N2 C3 H4 H5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.17512.45872.76162.67113.40563.83204.11164.61454.23222.45742.69742.63651.46852.0743
N21.17513.47463.60253.48764.49264.92445.23345.61735.34843.46793.51813.44682.64343.1538
C32.45873.47461.09411.09271.09213.09453.39074.11562.84182.56752.80573.49311.53572.1575
H42.76163.60251.09411.77401.77753.41493.99494.28593.02212.78982.58313.77102.16983.0665
H52.67113.48761.09271.77401.77174.10394.23945.15383.87043.51163.78014.31162.17702.5207
H63.40564.49261.09211.77751.77172.79632.95643.83542.29482.85203.22133.84162.17292.4747
C73.83204.92443.09453.41494.10392.79631.09511.09331.09281.52862.16862.15982.55902.7830
H84.11165.23343.39073.99494.23942.95641.09511.76981.77302.17253.07922.54982.77432.5495
H94.61455.61734.11564.28595.15383.83541.09331.76981.76482.16602.53982.45273.50013.7475
H104.23225.34842.84183.02213.87042.29481.09281.77301.76482.18462.49983.07812.88643.2394
C112.45743.46792.56752.78983.51162.85201.52862.17252.16602.18461.09641.09531.54092.1553
H122.69743.51812.80572.58313.78013.22132.16863.07922.53982.49981.09641.76032.15523.0532
H132.63653.44683.49313.77104.31163.84162.15982.54982.45273.07811.09531.76032.15312.4768
C141.46852.64341.53572.16982.17702.17292.55902.77433.50012.88641.54092.15522.15311.0979
H152.07433.15382.15753.06652.52072.47472.78302.54953.74753.23942.15533.05322.47681.0979

picture of Butanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C3 109.835 C1 C14 C11 109.468
C1 C14 H15 106.963 N2 C1 C14 178.581
C3 C14 C11 113.138 C3 C14 H15 108.882
H4 C3 H5 108.432 H4 C3 H6 108.789
H4 C3 C14 110.058 H5 C3 H6 108.373
H5 C3 C14 110.709 H6 C3 C14 110.420
C7 C11 H12 110.323 C7 C11 H13 109.699
C7 C11 C14 112.960 H8 C7 H9 107.941
H8 C7 H10 108.265 H8 C7 C11 110.714
H9 C7 H10 107.660 H9 C7 C11 110.298
H10 C7 C11 111.824 C11 C14 H15 108.356
H12 C11 H13 106.873 H12 C11 C14 108.441
H13 C11 C14 108.340
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability