Jump to
S1C2
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -187.550141 |
| Energy at 298.15K | -187.554172 |
| HF Energy | -186.927274 |
| Nuclear repulsion energy | 101.452837 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3668 |
3486 |
20.07 |
|
|
|
| 2 |
A |
3571 |
3393 |
3.13 |
|
|
|
| 3 |
A |
2381 |
2262 |
0.19 |
|
|
|
| 4 |
A |
1654 |
1571 |
15.03 |
|
|
|
| 5 |
A |
1222 |
1161 |
0.04 |
|
|
|
| 6 |
A |
820 |
779 |
16.66 |
|
|
|
| 7 |
A |
698 |
664 |
100.54 |
|
|
|
| 8 |
A |
379 |
360 |
0.95 |
|
|
|
| 9 |
A |
329 |
313 |
36.75 |
|
|
|
| 10 |
A |
208 |
197 |
14.88 |
|
|
|
| 11 |
B |
3668 |
3486 |
18.99 |
|
|
|
| 12 |
B |
3573 |
3395 |
11.93 |
|
|
|
| 13 |
B |
1653 |
1571 |
30.74 |
|
|
|
| 14 |
B |
1363 |
1295 |
85.99 |
|
|
|
| 15 |
B |
1221 |
1160 |
0.03 |
|
|
|
| 16 |
B |
754 |
716 |
350.65 |
|
|
|
| 17 |
B |
378 |
359 |
14.40 |
|
|
|
| 18 |
B |
208 |
198 |
18.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13873.4 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 13182.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.004 |
0.608 |
0.054 |
| C2 |
-0.004 |
-0.608 |
0.054 |
| N3 |
-0.004 |
1.962 |
-0.088 |
| N4 |
0.004 |
-1.962 |
-0.088 |
| H5 |
-0.317 |
2.459 |
0.733 |
| H6 |
0.863 |
2.342 |
-0.441 |
| H7 |
0.317 |
-2.459 |
0.733 |
| H8 |
-0.863 |
-2.342 |
-0.441 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2161 | 1.3616 | 2.5742 | 1.9976 | 1.9979 | 3.1570 | 3.1147 |
C2 | 1.2161 | | 2.5742 | 1.3616 | 3.1570 | 3.1147 | 1.9976 | 1.9979 | N3 | 1.3616 | 2.5742 | | 3.9244 | 1.0098 | 1.0100 | 4.5084 | 4.4038 | N4 | 2.5742 | 1.3616 | 3.9244 | | 4.5084 | 4.4038 | 1.0098 | 1.0100 | H5 | 1.9976 | 3.1570 | 1.0098 | 4.5084 | | 1.6685 | 4.9589 | 4.9730 | H6 | 1.9979 | 3.1147 | 1.0100 | 4.4038 | 1.6685 | | 4.9730 | 4.9928 | H7 | 3.1570 | 1.9976 | 4.5084 | 1.0098 | 4.9589 | 4.9730 | | 1.6685 | H8 | 3.1147 | 1.9979 | 4.4038 | 1.0100 | 4.9730 | 4.9928 | 1.6685 | |
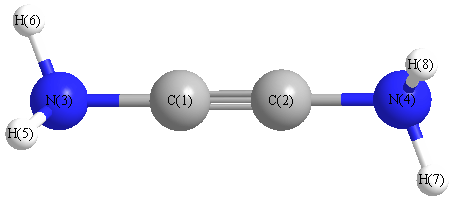 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
173.966 |
|
C1 |
N3 |
H5 |
113.957 |
| C1 |
N3 |
H6 |
113.973 |
|
C2 |
C1 |
N3 |
173.966 |
| C2 |
N4 |
H7 |
113.957 |
|
C2 |
N4 |
H8 |
113.973 |
| H5 |
N3 |
H6 |
111.404 |
|
H7 |
N4 |
H8 |
111.404 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -187.544116 |
| Energy at 298.15K | |
| HF Energy | -186.924775 |
| Nuclear repulsion energy | 101.992447 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3664 |
3482 |
0.00 |
|
|
|
| 2 |
A1 |
2394 |
2274 |
0.00 |
|
|
|
| 3 |
A1 |
1647 |
1565 |
0.00 |
|
|
|
| 4 |
A1 |
838 |
796 |
0.00 |
|
|
|
| 5 |
B1 |
495 |
470 |
0.00 |
|
|
|
| 6 |
B2 |
3669 |
3487 |
52.43 |
|
|
|
| 7 |
B2 |
1656 |
1573 |
58.89 |
|
|
|
| 8 |
B2 |
1436 |
1364 |
139.35 |
|
|
|
| 9 |
E |
3784 |
3596 |
45.66 |
|
|
|
| 9 |
E |
3784 |
3596 |
45.66 |
|
|
|
| 10 |
E |
1138 |
1081 |
1.76 |
|
|
|
| 10 |
E |
1138 |
1081 |
1.76 |
|
|
|
| 11 |
E |
412 |
391 |
1.20 |
|
|
|
| 11 |
E |
412 |
391 |
1.20 |
|
|
|
| 12 |
E |
192 |
182 |
27.73 |
|
|
|
| 12 |
E |
192 |
182 |
27.73 |
|
|
|
| 13 |
E |
551i |
524i |
262.42 |
|
|
|
| 13 |
E |
551i |
524i |
262.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12873.6 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 12232.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.609 |
| C2 |
0.000 |
0.000 |
-0.609 |
| N3 |
0.000 |
0.000 |
1.952 |
| N4 |
0.000 |
0.000 |
-1.952 |
| H5 |
0.000 |
0.860 |
2.468 |
| H6 |
0.000 |
-0.860 |
2.468 |
| H7 |
0.860 |
0.000 |
-2.468 |
| H8 |
-0.860 |
0.000 |
-2.468 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2187 | 1.3428 | 2.5615 | 2.0484 | 2.0484 | 3.1957 | 3.1957 |
C2 | 1.2187 | | 2.5615 | 1.3428 | 3.1957 | 3.1957 | 2.0484 | 2.0484 | N3 | 1.3428 | 2.5615 | | 3.9043 | 1.0033 | 1.0033 | 4.5035 | 4.5035 | N4 | 2.5615 | 1.3428 | 3.9043 | | 4.5035 | 4.5035 | 1.0033 | 1.0033 | H5 | 2.0484 | 3.1957 | 1.0033 | 4.5035 | | 1.7208 | 5.0844 | 5.0844 | H6 | 2.0484 | 3.1957 | 1.0033 | 4.5035 | 1.7208 | | 5.0844 | 5.0844 | H7 | 3.1957 | 2.0484 | 4.5035 | 1.0033 | 5.0844 | 5.0844 | | 1.7208 | H8 | 3.1957 | 2.0484 | 4.5035 | 1.0033 | 5.0844 | 5.0844 | 1.7208 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
120.962 |
| C1 |
N3 |
H6 |
120.962 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
120.962 |
|
C2 |
N4 |
H8 |
120.962 |
| H5 |
N3 |
H6 |
118.075 |
|
H7 |
N4 |
H8 |
118.075 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
