Jump to
S1C2
Vibrational Frequencies calculated at MP2/6-311G**
Geometric Data calculated at MP2/6-311G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -233.041107 |
| Energy at 298.15K | -233.052406 |
| HF Energy | -232.220603 |
| Nuclear repulsion energy | 199.776294 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3871 |
3679 |
16.24 |
|
|
|
| 2 |
A |
3177 |
3018 |
44.65 |
|
|
|
| 3 |
A |
3175 |
3017 |
30.72 |
|
|
|
| 4 |
A |
3170 |
3012 |
3.18 |
|
|
|
| 5 |
A |
3169 |
3011 |
2.64 |
|
|
|
| 6 |
A |
3152 |
2995 |
50.80 |
|
|
|
| 7 |
A |
3146 |
2989 |
2.47 |
|
|
|
| 8 |
A |
3077 |
2924 |
9.15 |
|
|
|
| 9 |
A |
3064 |
2911 |
24.97 |
|
|
|
| 10 |
A |
3060 |
2908 |
15.84 |
|
|
|
| 11 |
A |
1536 |
1459 |
9.46 |
|
|
|
| 12 |
A |
1526 |
1450 |
5.08 |
|
|
|
| 13 |
A |
1517 |
1441 |
2.95 |
|
|
|
| 14 |
A |
1504 |
1429 |
0.09 |
|
|
|
| 15 |
A |
1498 |
1423 |
0.00 |
|
|
|
| 16 |
A |
1490 |
1416 |
0.03 |
|
|
|
| 17 |
A |
1438 |
1367 |
4.13 |
|
|
|
| 18 |
A |
1424 |
1353 |
34.62 |
|
|
|
| 19 |
A |
1412 |
1341 |
20.40 |
|
|
|
| 20 |
A |
1384 |
1315 |
19.86 |
|
|
|
| 21 |
A |
1290 |
1226 |
18.28 |
|
|
|
| 22 |
A |
1271 |
1208 |
41.74 |
|
|
|
| 23 |
A |
1189 |
1130 |
58.00 |
|
|
|
| 24 |
A |
1060 |
1007 |
1.16 |
|
|
|
| 25 |
A |
1044 |
992 |
8.95 |
|
|
|
| 26 |
A |
968 |
920 |
0.01 |
|
|
|
| 27 |
A |
968 |
920 |
32.82 |
|
|
|
| 28 |
A |
949 |
902 |
1.20 |
|
|
|
| 29 |
A |
933 |
886 |
0.00 |
|
|
|
| 30 |
A |
770 |
732 |
2.25 |
|
|
|
| 31 |
A |
475 |
451 |
19.24 |
|
|
|
| 32 |
A |
467 |
444 |
10.38 |
|
|
|
| 33 |
A |
416 |
396 |
0.23 |
|
|
|
| 34 |
A |
347 |
330 |
23.50 |
|
|
|
| 35 |
A |
346 |
329 |
1.17 |
|
|
|
| 36 |
A |
318 |
302 |
71.59 |
|
|
|
| 37 |
A |
287 |
273 |
0.02 |
|
|
|
| 38 |
A |
272 |
259 |
4.82 |
|
|
|
| 39 |
A |
195 |
186 |
1.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30176.3 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 28673.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.006 |
-0.000 |
0.015 |
| H2 |
1.739 |
1.266 |
-0.216 |
| H3 |
0.202 |
2.146 |
-0.095 |
| H4 |
0.638 |
1.307 |
-1.605 |
| C5 |
0.683 |
1.258 |
-0.512 |
| H6 |
-1.969 |
0.887 |
0.070 |
| H7 |
-1.969 |
-0.888 |
0.069 |
| H8 |
-1.616 |
0.000 |
-1.433 |
| C9 |
-1.485 |
-0.000 |
-0.347 |
| H10 |
0.202 |
-2.146 |
-0.095 |
| H11 |
1.739 |
-1.265 |
-0.216 |
| H12 |
0.638 |
-1.308 |
-1.605 |
| C13 |
0.683 |
-1.258 |
-0.512 |
| H14 |
0.953 |
0.000 |
1.696 |
| O15 |
0.024 |
0.000 |
1.447 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
H4 |
C5 |
H6 |
H7 |
H8 |
C9 |
H10 |
H11 |
H12 |
C13 |
H14 |
O15 |
| C1 | | 2.1679 | 2.1593 | 2.1786 | 1.5281 | 2.1551 | 2.1551 | 2.1654 | 1.5227 | 2.1593 | 2.1679 | 2.1786 | 1.5281 | 1.9362 | 1.4324 |
H2 | 2.1679 | | 1.7756 | 1.7730 | 1.0964 | 3.7383 | 4.2974 | 3.7870 | 3.4661 | 3.7440 | 2.5310 | 3.1245 | 2.7513 | 2.4239 | 2.7028 | H3 | 2.1593 | 1.7756 | | 1.7817 | 1.0934 | 2.5153 | 3.7346 | 3.1154 | 2.7419 | 4.2929 | 3.7440 | 3.7948 | 3.4634 | 2.8946 | 2.6484 | H4 | 2.1786 | 1.7730 | 1.7817 | | 1.0946 | 3.1267 | 3.7970 | 2.6114 | 2.7924 | 3.7947 | 3.1243 | 2.6149 | 2.7886 | 3.5646 | 3.3759 | C5 | 1.5281 | 1.0964 | 1.0934 | 1.0946 | | 2.7408 | 3.4608 | 2.7783 | 2.5124 | 3.4634 | 2.7512 | 2.7887 | 2.5157 | 2.5560 | 2.4194 | H6 | 2.1551 | 3.7383 | 2.5153 | 3.1267 | 2.7408 | | 1.7751 | 1.7807 | 1.0935 | 3.7345 | 4.2974 | 3.7970 | 3.4608 | 3.4604 | 2.5804 | H7 | 2.1551 | 4.2974 | 3.7346 | 3.7970 | 3.4608 | 1.7751 | | 1.7807 | 1.0935 | 2.5152 | 3.7383 | 3.1265 | 2.7407 | 3.4605 | 2.5805 | H8 | 2.1654 | 3.7870 | 3.1154 | 2.6114 | 2.7783 | 1.7807 | 1.7807 | | 1.0939 | 3.1156 | 3.7870 | 2.6114 | 2.7784 | 4.0494 | 3.3145 | C9 | 1.5227 | 3.4661 | 2.7419 | 2.7924 | 2.5124 | 1.0935 | 1.0935 | 1.0939 | | 2.7420 | 3.4661 | 2.7924 | 2.5124 | 3.1816 | 2.3446 | H10 | 2.1593 | 3.7440 | 4.2929 | 3.7947 | 3.4634 | 3.7345 | 2.5152 | 3.1156 | 2.7420 | | 1.7755 | 1.7817 | 1.0934 | 2.8946 | 2.6484 | H11 | 2.1679 | 2.5310 | 3.7440 | 3.1243 | 2.7512 | 4.2974 | 3.7383 | 3.7870 | 3.4661 | 1.7755 | | 1.7730 | 1.0964 | 2.4241 | 2.7029 | H12 | 2.1786 | 3.1245 | 3.7948 | 2.6149 | 2.7887 | 3.7970 | 3.1265 | 2.6114 | 2.7924 | 1.7817 | 1.7730 | | 1.0946 | 3.5647 | 3.3759 | C13 | 1.5281 | 2.7513 | 3.4634 | 2.7886 | 2.5157 | 3.4608 | 2.7407 | 2.7784 | 2.5124 | 1.0934 | 1.0964 | 1.0946 | | 2.5560 | 2.4194 | H14 | 1.9362 | 2.4239 | 2.8946 | 3.5646 | 2.5560 | 3.4604 | 3.4605 | 4.0494 | 3.1816 | 2.8946 | 2.4241 | 3.5647 | 2.5560 | | 0.9618 | O15 | 1.4324 | 2.7028 | 2.6484 | 3.3759 | 2.4194 | 2.5804 | 2.5805 | 3.3145 | 2.3446 | 2.6484 | 2.7029 | 3.3759 | 2.4194 | 0.9618 | |
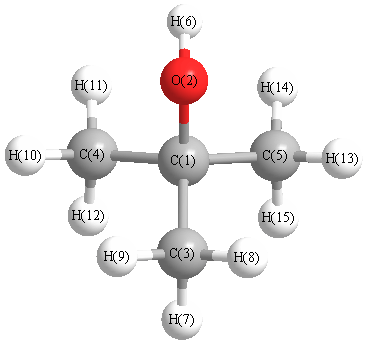 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
H2 |
110.302 |
|
C1 |
C5 |
H3 |
109.796 |
| C1 |
C5 |
H4 |
111.257 |
|
C1 |
C9 |
H6 |
109.835 |
| C1 |
C9 |
H7 |
109.836 |
|
C1 |
C9 |
H8 |
110.632 |
| C1 |
C13 |
H10 |
109.795 |
|
C1 |
C13 |
H11 |
110.303 |
| C1 |
C13 |
H12 |
111.258 |
|
C1 |
O15 |
H14 |
106.271 |
| H2 |
C5 |
H3 |
108.350 |
|
H2 |
C5 |
H4 |
108.036 |
| H3 |
C5 |
H4 |
109.031 |
|
C5 |
C1 |
C9 |
110.877 |
| C5 |
C1 |
C13 |
110.801 |
|
C5 |
C1 |
O15 |
109.575 |
| H6 |
C9 |
H7 |
108.517 |
|
H6 |
C9 |
H8 |
108.990 |
| H7 |
C9 |
H8 |
108.991 |
|
C9 |
C1 |
C13 |
110.877 |
| C9 |
C1 |
O15 |
104.969 |
|
H10 |
C13 |
H11 |
108.348 |
| H10 |
C13 |
H12 |
109.029 |
|
H11 |
C13 |
H12 |
108.036 |
| C13 |
C1 |
O15 |
109.575 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
