return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9NO (Butanamide)

using model chemistry: MP2/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G**
 hartrees
Energy at 0K-286.976705
Energy at 298.15K-286.987136
HF Energy-286.062776
Nuclear repulsion energy238.873753
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3833 3589 40.52      
2 A 3683 3449 46.00      
3 A 3222 3017 29.46      
4 A 3221 3017 45.50      
5 A 3201 2998 0.29      
6 A 3164 2963 13.87      
7 A 3145 2945 18.91      
8 A 3123 2924 23.77      
9 A 3111 2914 13.98      
10 A 1815 1700 234.55      
11 A 1663 1557 105.83      
12 A 1566 1467 3.86      
13 A 1561 1462 5.86      
14 A 1549 1451 2.59      
15 A 1523 1426 5.89      
16 A 1469 1375 20.35      
17 A 1462 1369 46.31      
18 A 1400 1311 42.26      
19 A 1360 1274 23.05      
20 A 1298 1216 47.32      
21 A 1293 1211 7.75      
22 A 1171 1096 0.56      
23 A 1151 1078 1.17      
24 A 1106 1036 0.49      
25 A 1088 1019 3.63      
26 A 961 900 2.44      
27 A 909 851 0.59      
28 A 875 819 2.75      
29 A 767 718 4.92      
30 A 674 631 16.66      
31 A 600 562 9.11      
32 A 523 490 20.84      
33 A 436 408 4.36      
34 A 351 328 2.30      
35 A 258 242 0.55      
36 A 238 223 247.80      
37 A 189 177 6.94      
38 A 96 90 1.64      
39 A 28 26 0.91      

Unscaled Zero Point Vibrational Energy (zpe) 29540.3 cm-1
Scaled (by 0.9365) Zero Point Vibrational Energy (zpe) 27664.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G**
ABC
0.28767 0.06148 0.05269

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.721 -0.211 0.025
H2 -2.863 -0.436 1.082
H3 -3.511 0.473 -0.281
H4 -2.848 -1.140 -0.533
C5 -1.344 0.395 -0.222
H6 -1.230 0.645 -1.278
H7 -1.231 1.329 0.327
C8 -0.218 -0.543 0.192
H9 -0.261 -1.472 -0.382
H10 -0.334 -0.825 1.243
N11 2.193 -0.778 -0.078
H12 3.103 -0.387 -0.255
H13 2.037 -1.741 -0.316
C14 1.150 0.102 0.043
O15 1.323 1.319 0.089

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 N11 H12 H13 C14 O15
C11.09031.08921.09051.52362.15692.16362.53012.79372.74894.94745.83295.00893.88354.3235
H21.09031.76171.76162.16723.06702.51972.79323.16082.56425.19926.11475.26044.18084.6465
H31.08921.76171.76142.16892.49542.51003.47863.78893.75505.84336.66965.97334.68724.9211
H41.09051.76161.76142.17072.52163.07352.79282.61263.09415.07476.00504.92674.22624.8812
C51.52362.16722.16892.17071.09161.08921.52292.16442.15713.72974.51564.00012.52562.8397
H62.15693.06702.49542.52161.09161.74442.14392.49463.05253.89654.56974.15772.77562.9727
H72.16362.51972.51003.07351.08921.74442.13263.04802.50644.04094.69744.52932.69372.5650
C82.53012.79323.47862.79281.52292.14392.13261.09361.09422.43753.35462.60331.52002.4187
H92.79373.16083.78892.61262.16442.49463.04801.09361.75132.56883.53742.31472.15693.2436
H102.74892.56423.75503.09412.15713.05252.50641.09421.75132.85193.77502.98192.12202.9449
N114.94745.19925.84335.07473.72973.89654.04092.43752.56882.85191.00641.00381.37012.2765
H125.83296.11476.66966.00504.51564.56974.69743.35463.53743.77501.00641.72452.03492.4890
H135.00895.26045.97334.92674.00014.15774.52932.60332.31472.98191.00381.72452.07623.1675
C143.88354.18084.68724.22622.52562.77562.69371.52002.15692.12201.37012.03492.07621.2296
O154.32354.64654.92114.88122.83972.97272.56502.41873.24362.94492.27652.48903.16751.2296

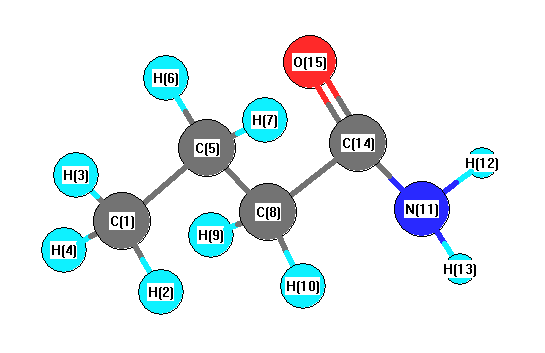
picture of Butanamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.027 C1 C5 H7 110.702
C1 C5 C8 112.297 H2 C1 H3 107.859
H2 C1 H4 107.761 H2 C1 C5 110.923
H3 C1 H4 107.827 H3 C1 C5 111.125
H4 C1 C5 111.191 C5 C8 H9 110.557
C5 C8 H10 109.940 C5 C8 C14 112.202
H6 C5 H7 106.242 H6 C5 C8 109.054
H7 C5 C8 108.315 C8 C14 N11 114.907
C8 C14 O15 122.856 H9 C8 H10 106.354
H9 C8 C14 110.164 H10 C8 C14 107.409
N11 C14 O15 122.153 H12 N11 H13 118.162
H12 N11 C14 116.975 H13 N11 C14 121.227
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability