return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5N ((Z)-2-Butenenitrile)

using model chemistry: MP2/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/TZVP
 hartrees
Energy at 0K-209.595971
Energy at 298.15K-209.600391
HF Energy-208.871106
Nuclear repulsion energy141.772378
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3247 3078 3.39      
2 A 3214 3047 7.99      
3 A 3191 3024 2.79      
4 A 3085 2924 9.28      
5 A 2196 2082 0.38      
6 A 1696 1608 2.32      
7 A 1510 1432 12.65      
8 A 1449 1374 4.53      
9 A 1414 1340 2.41      
10 A 1274 1208 0.26      
11 A 1135 1076 0.13      
12 A 975 924 10.84      
13 A 915 867 1.89      
14 A 659 625 1.15      
15 A 390 370 0.09      
16 A 157 149 4.19      
17 A 3159 2994 8.51      
18 A 1494 1417 8.58      
19 A 1070 1015 3.05      
20 A 935 886 0.01      
21 A 719 682 39.89      
22 A 468 443 1.63      
23 A 259 245 2.48      
24 A 138 131 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 17374.7 cm-1
Scaled (by 0.9479) Zero Point Vibrational Energy (zpe) 16469.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/TZVP
ABC
0.39727 0.11674 0.09176

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/TZVP

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.175 0.291 0.000
N2 -2.183 -0.294 0.000
C3 0.000 0.981 0.000
H4 -0.035 2.038 0.000
C5 1.211 0.379 0.000
H6 2.067 1.001 0.000
C7 1.427 -1.083 0.000
H8 0.485 -1.627 0.000
H9 1.992 -1.380 0.883
H10 1.992 -1.380 -0.883

Atom - Atom Distances (Å)
  C1 N2 C3 H4 C5 H6 C7 H8 H9 H10
C11.16631.36182.08522.38753.31782.94232.53663.68793.6879
N21.16632.52813.17063.46074.44273.69552.98254.40374.4037
C31.36182.52811.05751.35272.06662.50892.65253.21253.2125
H42.08523.17061.05752.07462.34283.44583.70144.07024.0702
C52.38753.46071.35272.07461.05761.47752.13372.11712.1171
H63.31784.44272.06662.34281.05762.17993.06772.54072.5407
C72.94233.69552.50893.44581.47752.17991.08841.08961.0896
H82.53662.98252.65253.70142.13373.06771.08841.76451.7645
H93.68794.40373.21254.07022.11712.54071.08961.76451.7661
H103.68794.40373.21254.07022.11712.54071.08961.76451.7661

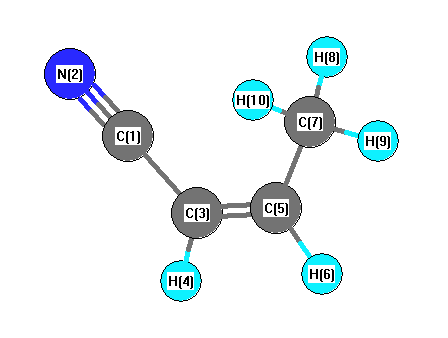
picture of (Z)-2-Butenenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H4 118.527 C1 C3 C5 123.182
N2 C1 C3 179.732 C3 C5 H6 117.534
C3 C5 C7 124.812 H4 C3 C5 118.291
C5 C7 H8 111.619 C5 C7 H9 110.207
C5 C7 H10 110.207 H6 C5 C7 117.655
H8 C7 H9 108.217 H8 C7 H10 108.217
H9 C7 H10 108.275
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability