return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: MP2/Def2TZVPP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/Def2TZVPP
 hartrees
Energy at 0K-398.311334
Energy at 298.15K-398.321342
HF Energy-396.895864
Nuclear repulsion energy326.843231
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/Def2TZVPP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3822 3637 62.96      
2 A 3764 3582 91.38      
3 A 3630 3454 9.58      
4 A 3530 3359 3.60      
5 A 3173 3019 13.29      
6 A 3120 2968 8.30      
7 A 3057 2909 36.21      
8 A 1792 1705 225.87      
9 A 1656 1575 32.05      
10 A 1514 1441 4.59      
11 A 1435 1366 24.88      
12 A 1419 1350 25.23      
13 A 1400 1332 48.45      
14 A 1386 1318 4.92      
15 A 1321 1257 13.28      
16 A 1255 1194 13.30      
17 A 1194 1136 2.16      
18 A 1161 1105 204.53      
19 A 1147 1091 43.64      
20 A 1107 1053 94.71      
21 A 1043 992 47.56      
22 A 1006 957 1.92      
23 A 881 838 153.29      
24 A 823 783 28.35      
25 A 754 717 19.03      
26 A 626 596 82.91      
27 A 597 568 22.32      
28 A 524 499 161.68      
29 A 505 481 29.57      
30 A 457 435 8.53      
31 A 290 276 7.45      
32 A 285 271 31.11      
33 A 271 257 6.73      
34 A 208 198 1.59      
35 A 162 154 4.55      
36 A 43 41 1.25      

Unscaled Zero Point Vibrational Energy (zpe) 25176.7 cm-1
Scaled (by 0.9515) Zero Point Vibrational Energy (zpe) 23955.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/Def2TZVPP
ABC
0.11735 0.07839 0.05356

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/Def2TZVPP

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.779 -0.535 0.014
O2 2.048 -0.345 -0.389
O3 0.403 -1.537 0.581
C4 -0.087 0.672 -0.311
C5 -1.504 0.433 0.187
O6 -2.093 -0.702 -0.399
N7 0.421 1.920 0.234
H8 2.539 -1.142 -0.150
H9 -0.120 0.755 -1.398
H10 -1.489 0.368 1.280
H11 -2.101 1.297 -0.089
H12 -1.625 -1.464 -0.048
H13 1.325 2.143 -0.157
H14 0.532 1.854 1.237

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.34571.21091.52042.48502.90632.49111.86902.11352.74913.41482.57742.73902.6952
O21.34572.25162.36643.68144.15712.85780.96622.63243.97544.47273.85502.60143.1273
O31.21092.25162.43202.76962.80893.47492.29153.07332.77383.84082.12423.86533.4563
C41.52042.36642.43201.52042.43321.45403.19551.09112.14192.12062.64502.04552.0435
C52.48503.68142.76961.52041.40692.43324.35132.12891.09501.08631.91533.32382.6951
O62.90634.15712.80892.43321.40693.68804.65952.64882.08082.02300.96124.45454.0125
N72.49112.85783.47491.45402.43323.68803.74312.07722.67462.61843.96501.00981.0113
H81.86900.96622.29153.19554.35134.65953.74313.49664.53245.24234.17703.50243.8639
H92.11352.63243.07331.09112.12892.64882.07723.49663.03272.43613.00232.35742.9285
H102.74913.97542.77382.14191.09502.08082.67464.53243.03271.76432.26643.62442.5084
H113.41484.47273.84082.12061.08632.02302.61845.24232.43611.76432.80223.53032.9998
H122.57743.85502.12422.64501.91530.96123.96504.17703.00232.26642.80224.66144.1605
H132.73902.60143.86532.04553.32384.45451.00983.50242.35743.62443.53034.66141.6298
H142.69523.12733.45632.04352.69514.01251.01133.86392.92852.50842.99984.16051.6298

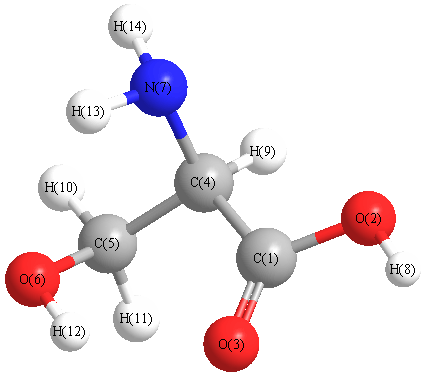
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 106.730 C1 C4 C5 109.613
C1 C4 N7 113.737 C1 C4 H9 106.901
O2 C1 O3 123.364 O2 C1 C4 111.156
O3 C1 C4 125.469 C4 C5 O6 112.386
C4 C5 H10 108.874 C4 C5 H11 107.718
C4 N7 H13 110.958 C4 N7 H14 110.686
C5 C4 N7 109.759 C5 C4 H9 108.085
C5 O6 H12 106.427 O6 C5 H10 111.942
O6 C5 H11 107.772 N7 C4 H9 108.560
H10 C5 H11 107.963 H13 N7 H14 107.492
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability