return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2(SH)CH(CH3)CH3 (1-Propanethiol, 2-methyl-)

using model chemistry: MP2/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/cc-pVDZ
 hartrees
Energy at 0K-555.557738
Energy at 298.15K-555.568481
HF Energy-554.836500
Nuclear repulsion energy233.759702
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3185 3034 19.15      
2 A 3175 3024 26.10      
3 A 3169 3019 47.67      
4 A 3165 3015 4.47      
5 A 3153 3003 8.26      
6 A 3086 2939 32.08      
7 A 3077 2931 8.62      
8 A 3074 2928 14.90      
9 A 3070 2924 20.23      
10 A 2776 2644 6.14      
11 A 1506 1434 10.44      
12 A 1500 1429 7.08      
13 A 1491 1420 2.77      
14 A 1483 1412 0.48      
15 A 1473 1403 3.44      
16 A 1416 1348 6.31      
17 A 1401 1335 5.06      
18 A 1374 1309 1.63      
19 A 1359 1295 1.22      
20 A 1276 1215 20.07      
21 A 1242 1183 4.95      
22 A 1209 1151 3.58      
23 A 1165 1110 2.42      
24 A 1094 1042 2.65      
25 A 997 950 1.41      
26 A 980 934 0.74      
27 A 932 888 1.34      
28 A 916 872 1.95      
29 A 885 843 1.68      
30 A 825 786 2.35      
31 A 752 717 0.60      
32 A 427 407 0.04      
33 A 396 377 0.88      
34 A 340 324 0.08      
35 A 264 251 0.02      
36 A 233 222 0.32      
37 A 211 201 1.73      
38 A 170 162 13.56      
39 A 83 79 8.23      

Unscaled Zero Point Vibrational Energy (zpe) 29164.5 cm-1
Scaled (by 0.9525) Zero Point Vibrational Energy (zpe) 27779.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/cc-pVDZ
ABC
0.24607 0.07137 0.05962

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.992 1.450 0.168
H2 0.088 2.041 -0.051
H3 1.151 1.466 1.262
H4 1.852 1.953 -0.307
S5 -1.901 0.064 -0.127
H6 -2.689 -0.854 0.468
C7 2.165 -0.770 -0.060
H8 2.104 -1.803 -0.442
H9 3.025 -0.277 -0.544
H10 2.370 -0.817 1.025
C11 -0.306 -0.729 0.326
H12 -0.322 -1.779 -0.012
H13 -0.184 -0.718 1.424
C14 0.870 0.006 -0.329
H15 0.691 0.021 -1.422

Atom - Atom Distances (Å)
  C1 H2 H3 H4 S5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.10181.10521.10323.22204.35372.52123.49182.76082.78792.54213.49132.76841.53242.1590
H21.10181.78401.78462.80564.04513.49484.35803.77353.81172.82323.84193.14032.19772.5146
H31.10521.78401.78573.63434.55602.78793.80713.13192.59842.79573.78422.56452.17693.0820
H41.10321.78461.78574.20565.39472.75153.76662.53033.11643.50034.32903.77822.18052.5147
S53.22202.80563.63434.20561.34824.15134.43014.95544.51021.83772.42912.44272.77922.8977
H64.35374.04514.55605.39471.34824.88414.97085.83215.08992.39062.58662.68533.74783.9704
C72.52123.49482.78792.75154.15134.88411.10321.10311.10492.50192.68492.77921.53322.1568
H83.49184.35803.80713.76664.43014.97081.10321.78551.78712.74832.46443.14552.19262.5065
H92.76083.77353.13192.53034.95545.83211.10311.78551.78403.47283.70733.79032.18382.5110
H102.78793.81172.59843.11644.51025.08991.10491.78711.78402.76723.04112.58652.18123.0828
C112.54212.82322.79573.50031.83772.39062.50192.74833.47282.76721.10301.10511.53422.1474
H123.49133.84193.78424.32902.42912.58662.68492.46443.70733.04111.10301.79122.16982.5003
H132.76843.14032.56453.77822.44272.68532.77923.14553.79032.58651.10511.79122.16993.0676
C141.53242.19772.17692.18052.77923.74781.53322.19262.18382.18121.53422.16982.16991.1075
H152.15902.51463.08202.51472.89773.97042.15682.50652.51103.08282.14742.50033.06761.1075

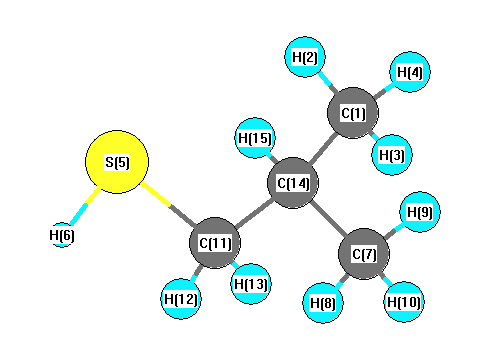
picture of 1-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C7 110.658 C1 C14 C11 111.986
C1 C14 H15 108.673 H2 C1 H3 107.871
H2 C1 H4 108.070 H2 C1 C14 112.049
H3 C1 H4 107.920 H3 C1 C14 110.194
H4 C1 C14 110.597 S5 C11 H12 108.807
S5 C11 H13 109.689 S5 C11 C14 110.705
H6 S5 C11 96.030 C7 C14 C11 109.300
C7 C14 H15 108.444 H8 C7 H9 108.049
H8 C7 H10 108.058 H8 C7 C14 111.496
H9 C7 H10 107.803 H9 C7 C14 110.808
H10 C7 C14 110.493 C11 C14 H15 107.669
H12 C11 H13 108.424 H12 C11 C14 109.644
H13 C11 C14 109.533
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability