return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

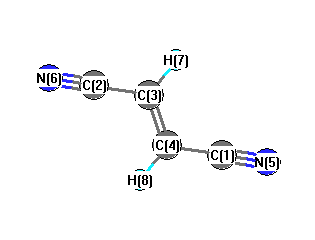
All results from a given calculation for C4H2N2 (Fumaronitrile)

using model chemistry: MP4=FULL/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP4=FULL/aug-cc-pVDZ
 hartrees
Energy at 0K-262.449657
Energy at 298.15K-262.450606
HF Energy-261.511652
Nuclear repulsion energy159.566865
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3205 3126        
2 Ag 2124 2072        
3 Ag 1631 1590        
4 Ag 1304 1272        
5 Ag 1003 978        
6 Ag 513 501        
7 Ag 239 233        
8 Au 953 929        
9 Au 520 507        
10 Au 122 119        
11 Bg 818 797        
12 Bg 354 346        
13 Bu 3210 3131        
14 Bu 2143 2089        
15 Bu 1269 1237        
16 Bu 999 975        
17 Bu 503 490        
18 Bu 128 125        

Unscaled Zero Point Vibrational Energy (zpe) 10517.6 cm-1
Scaled (by 0.9752) Zero Point Vibrational Energy (zpe) 10256.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/aug-cc-pVDZ
ABC
1.49396 0.04808 0.04658

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/aug-cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.344 0.589 0.000
C2 0.344 -0.589 0.000
C3 0.344 1.859 0.000
C4 -0.344 -1.859 0.000
N5 0.893 2.913 0.000
N6 -0.893 -2.913 0.000
H7 -1.439 0.609 0.000
H8 1.439 -0.609 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 N6 H7 H8
C11.36361.44452.44802.63323.54491.09502.1476
C21.36362.44801.44453.54492.63322.14761.0950
C31.44452.44803.78151.18874.93022.17692.7004
C42.44801.44453.78154.93021.18872.70042.1769
N52.63323.54491.18874.93026.09423.27813.5645
N63.54492.63324.93021.18876.09423.56453.2781
H71.09502.14762.17692.70043.27813.56453.1245
H82.14761.09502.70042.17693.56453.27813.1245

picture of Fumaronitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 121.301 C1 C2 H8 121.354
C1 C3 N5 179.122 C2 C1 C3 121.301
C2 C1 H7 121.354 C2 C4 N6 179.122
C3 C1 H7 117.345 C4 C2 H8 117.345
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability