return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Vinyl sulfoxide)

using model chemistry: QCISD/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-311G**
 hartrees
Energy at 0K-628.248227
Energy at 298.15K-628.254488
Nuclear repulsion energy289.816568
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3266 3116 0.57      
2 A' 3207 3060 7.35      
3 A' 3164 3019 2.53      
4 A' 1674 1597 1.26      
5 A' 1421 1356 15.25      
6 A' 1289 1230 0.83      
7 A' 1070 1021 71.79      
8 A' 1012 965 19.76      
9 A' 997 951 43.21      
10 A' 944 900 11.63      
11 A' 708 675 70.53      
12 A' 645 615 5.15      
13 A' 510 486 1.70      
14 A' 305 291 3.12      
15 A' 224 214 2.77      
16 A' 99 95 0.73      
17 A" 3265 3116 1.28      
18 A" 3203 3056 3.98      
19 A" 3164 3018 4.58      
20 A" 1665 1589 2.38      
21 A" 1417 1352 3.26      
22 A" 1273 1215 9.40      
23 A" 1005 959 1.33      
24 A" 987 942 47.24      
25 A" 938 895 5.12      
26 A" 661 631 14.87      
27 A" 560 534 5.56      
28 A" 484 461 9.99      
29 A" 268 256 8.40      
30 A" 156 149 0.66      

Unscaled Zero Point Vibrational Energy (zpe) 19788.9 cm-1
Scaled (by 0.9541) Zero Point Vibrational Energy (zpe) 18880.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-311G**
ABC
0.15828 0.07630 0.06829

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-311G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.588 -0.565 0.000
O2 1.337 0.741 0.000
C3 -0.628 -0.468 1.329
C4 -0.628 -0.468 -1.329
C5 -0.628 0.600 2.129
C6 -0.628 0.600 -2.129
H7 -1.279 -1.333 1.433
H8 -1.279 -1.333 -1.433
H9 -1.321 0.682 2.961
H10 -1.321 0.682 -2.961
H11 0.076 1.408 1.946
H12 0.076 1.408 -1.946

Atom - Atom Distances (Å)
  S1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
S11.50521.80381.80382.71432.71432.47572.47573.73713.73712.81802.8180
O21.50522.66242.66242.90082.90083.63293.63293.97973.97972.41302.4130
C31.80382.66242.65841.33423.61941.08772.96722.11324.49562.09653.8394
C41.80382.66242.65843.61941.33422.96721.08774.49562.11323.83942.0965
C52.71432.90081.33423.61944.25832.15494.10511.08605.13811.08744.2136
C62.71432.90083.61941.33424.25834.10512.15495.13811.08604.21361.0874
H72.47573.63291.08772.96722.15494.10512.86702.52884.83483.10034.5574
H82.47573.63292.96721.08774.10512.15492.86704.83482.52884.55743.1003
H93.73713.97972.11324.49561.08605.13812.52884.83485.92281.87355.1536
H103.73713.97974.49562.11325.13811.08604.83482.52885.92285.15361.8735
H112.81802.41302.09653.83941.08744.21363.10034.55741.87355.15363.8917
H122.81802.41303.83942.09654.21361.08744.55743.10035.15361.87353.8917

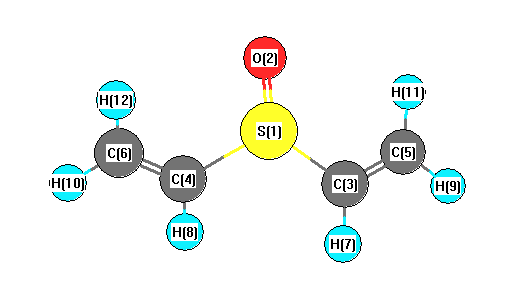
picture of Vinyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 119.002 S1 C3 H7 115.546
S1 C4 C6 119.002 S1 C4 H8 115.546
O2 S1 C3 106.793 O2 S1 C4 106.793
C3 S1 C4 94.929 C3 C5 H9 121.316
C3 C5 H11 119.593 C4 C6 H10 121.316
C4 C6 H12 119.593 C5 C3 H7 125.383
C6 C4 H8 125.383 H9 C5 H11 119.091
H10 C6 H12 119.091
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability