return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: QCISD/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-311G**
 hartrees
Energy at 0K-398.079807
Energy at 298.15K-398.089959
HF Energy-396.833832
Nuclear repulsion energy325.703849
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3892 3713 61.21      
2 A 3841 3665 74.11      
3 A 3616 3450 2.59      
4 A 3529 3367 0.88      
5 A 3143 2999 21.94      
6 A 3082 2941 14.12      
7 A 3020 2882 50.62      
8 A 1832 1748 239.32      
9 A 1679 1601 33.29      
10 A 1535 1464 3.00      
11 A 1478 1410 45.86      
12 A 1442 1376 7.59      
13 A 1426 1361 55.24      
14 A 1410 1345 11.20      
15 A 1351 1289 6.04      
16 A 1268 1210 16.03      
17 A 1224 1168 21.81      
18 A 1205 1150 202.47      
19 A 1170 1116 26.53      
20 A 1129 1078 92.00      
21 A 1061 1012 38.15      
22 A 1021 975 2.01      
23 A 935 892 169.33      
24 A 832 794 17.06      
25 A 752 717 27.32      
26 A 629 600 100.30      
27 A 585 558 11.26      
28 A 548 523 203.11      
29 A 526 502 9.31      
30 A 447 427 7.76      
31 A 313 298 7.47      
32 A 303 289 16.24      
33 A 264 252 23.19      
34 A 223 213 3.51      
35 A 173 165 2.95      
36 A 39 37 0.84      

Unscaled Zero Point Vibrational Energy (zpe) 25460.9 cm-1
Scaled (by 0.9541) Zero Point Vibrational Energy (zpe) 24292.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-311G**
ABC
0.11967 0.07816 0.05123

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-311G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.785 -0.543 0.006
O2 2.060 -0.335 -0.370
O3 0.413 -1.555 0.554
C4 -0.091 0.671 -0.310
C5 -1.516 0.436 0.197
O6 -2.116 -0.694 -0.391
N7 0.432 1.924 0.227
H8 2.544 -1.134 -0.134
H9 -0.132 0.754 -1.403
H10 -1.495 0.365 1.297
H11 -2.114 1.310 -0.074
H12 -1.648 -1.450 -0.033
H13 1.336 2.137 -0.178
H14 0.561 1.847 1.231

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.34561.20911.52992.50782.93182.50171.86152.12332.77273.44112.59672.74182.6954
O21.34562.24792.37573.70224.19142.84810.96342.65673.98844.49603.88652.58333.0949
O31.20912.24792.43992.79482.83363.49372.27883.07532.80683.87102.14553.87453.4719
C41.52992.37572.43991.53072.44341.46033.19921.09732.15502.13412.64562.04992.0454
C52.50783.70222.79481.53071.40782.45194.36612.13971.10201.09311.90453.34182.7158
O62.93184.19142.83362.44341.40783.70544.68812.65702.08702.02900.95844.46924.0317
N72.50172.84813.49371.46032.45193.70543.73412.08472.69912.63613.97231.01291.0147
H81.86150.96342.27883.19924.36614.68813.73413.51254.54005.26064.20543.48723.8327
H92.12332.65673.07531.09732.13972.65702.08473.51253.04922.44993.00582.35962.9351
H102.77273.98842.80682.15501.10202.08702.69914.54003.04921.77612.25563.65012.5349
H113.44114.49603.87102.13411.09312.02902.63615.26062.44991.77612.79953.54863.0240
H122.59673.88652.14552.64561.90450.95843.97234.20543.00582.25562.79954.66794.1653
H132.74182.58333.87452.04993.34184.46921.01293.48722.35963.65013.54864.66791.6337
H142.69543.09493.47192.04542.71584.03171.01473.83272.93512.53493.02404.16531.6337

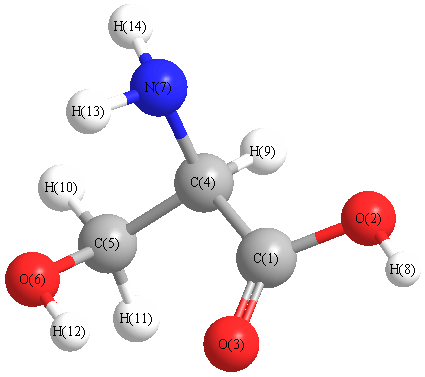
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 106.269 C1 C4 C5 110.050
C1 C4 N7 113.552 C1 C4 H9 106.685
O2 C1 O3 123.173 O2 C1 C4 111.256
O3 C1 C4 125.544 C4 C5 O6 112.439
C4 C5 H10 108.794 C4 C5 H11 107.682
C4 N7 H13 110.659 C4 N7 H14 110.162
C5 C4 N7 110.097 C5 C4 H9 107.881
C5 O6 H12 105.615 O6 C5 H10 111.939
O6 C5 H11 107.784 N7 C4 H9 108.355
H10 C5 H11 108.019 H13 N7 H14 107.374
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability