return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NHNHCH3 (dimethyl hydrazine)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-189.933124
Energy at 298.15K-189.943608
Nuclear repulsion energy136.799522
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3588 3378 0.60      
2 A 3190 3003 2.87      
3 A 3149 2965 15.10      
4 A 3075 2895 90.47      
5 A 1586 1493 2.43      
6 A 1577 1484 0.10      
7 A 1550 1459 0.26      
8 A 1502 1414 0.17      
9 A 1300 1224 17.35      
10 A 1190 1120 0.11      
11 A 1166 1097 0.73      
12 A 963 907 27.99      
13 A 770 725 132.78      
14 A 346 326 8.00      
15 A 331 312 0.71      
16 A 187 176 1.02      
17 B 3604 3393 0.98      
18 B 3190 3003 58.04      
19 B 3148 2963 74.47      
20 B 3061 2881 7.07      
21 B 1574 1482 8.66      
22 B 1543 1453 11.21      
23 B 1495 1408 1.64      
24 B 1481 1394 0.26      
25 B 1207 1137 1.93      
26 B 1168 1099 25.09      
27 B 1049 988 0.08      
28 B 820 772 35.07      
29 B 519 488 2.23      
30 B 234 221 0.94      

Unscaled Zero Point Vibrational Energy (zpe) 24780.6 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 23328.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.49169 0.18722 0.15352

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.160 0.697 -0.691
N2 0.160 -0.697 -0.691
C3 0.160 1.433 0.533
C4 -0.160 -1.433 0.533
H5 -1.154 0.758 -0.879
H6 1.154 -0.758 -0.879
H7 -0.111 2.477 0.372
H8 -0.347 1.084 1.440
H9 1.237 1.386 0.704
H10 0.111 -2.477 0.372
H11 0.347 -1.084 1.440
H12 -1.237 -1.386 0.704

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
N11.43011.46372.45691.01381.96922.07402.17372.09023.35842.82272.7281
N21.43012.45691.46371.96921.01383.35842.82272.72812.07402.17372.0902
C31.46372.45692.88452.04382.79011.09021.09611.09153.91432.68203.1506
C42.45691.46372.88452.79012.04383.91432.68203.15061.09021.09611.0915
H51.01381.96922.04382.79012.76112.36882.47712.93533.69203.32002.6659
H61.96921.01382.79012.04382.76113.69203.32002.66592.36882.47712.9353
H72.07403.35841.09023.91432.36883.69201.77151.76544.95943.74564.0374
H82.17372.82271.09612.68202.47713.32001.77151.77243.74562.27572.7261
H92.09022.72811.09153.15062.93532.66591.76541.77244.03742.72613.7147
H103.35842.07403.91431.09023.69202.36884.95943.74564.03741.77151.7654
H112.82272.17372.68201.09613.32002.47713.74562.27572.72611.77151.7724
H122.72812.09023.15061.09152.66592.93534.03742.72613.71471.76541.7724

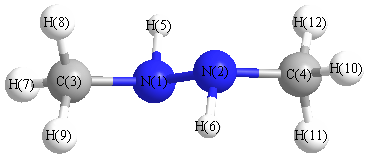
picture of dimethyl hydrazine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 116.211 N1 N2 H6 106.108
N1 C3 H7 107.709 N1 C3 H8 115.496
N1 C3 H9 108.908 N2 N1 C3 116.211
N2 N1 H5 106.108 N2 C4 H10 107.709
N2 C4 H11 115.496 N2 C4 H12 108.908
C3 N1 H5 109.833 C4 N2 H6 109.833
H7 C3 H8 108.243 H7 C3 H9 108.031
H8 C3 H9 108.230 H10 C4 H11 108.243
H10 C4 H12 108.031 H11 C4 H12 108.230
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability