Jump to
S1C2
Energy calculated at QCISD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -341.510568 |
| Energy at 298.15K | |
| HF Energy | -340.537494 |
| Nuclear repulsion energy | 244.488644 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3134 |
2963 |
58.82 |
|
|
|
| 2 |
A1 |
1930 |
1825 |
645.15 |
|
|
|
| 3 |
A1 |
1601 |
1514 |
4.15 |
|
|
|
| 4 |
A1 |
1438 |
1360 |
2.31 |
|
|
|
| 5 |
A1 |
1233 |
1166 |
246.43 |
|
|
|
| 6 |
A1 |
979 |
925 |
16.34 |
|
|
|
| 7 |
A1 |
829 |
783 |
42.72 |
|
|
|
| 8 |
A1 |
725 |
685 |
0.42 |
|
|
|
| 9 |
A2 |
3168 |
2995 |
0.00 |
|
|
|
| 10 |
A2 |
1275 |
1206 |
0.00 |
|
|
|
| 11 |
A2 |
1196 |
1131 |
0.00 |
|
|
|
| 12 |
A2 |
202i |
191i |
0.00 |
|
|
|
| 13 |
B1 |
3189 |
3014 |
39.96 |
|
|
|
| 14 |
B1 |
1288 |
1217 |
2.73 |
|
|
|
| 15 |
B1 |
879 |
831 |
1.25 |
|
|
|
| 16 |
B1 |
751 |
710 |
28.16 |
|
|
|
| 17 |
B1 |
122 |
116 |
2.78 |
|
|
|
| 18 |
B2 |
3126 |
2955 |
22.05 |
|
|
|
| 19 |
B2 |
1587 |
1500 |
2.31 |
|
|
|
| 20 |
B2 |
1443 |
1364 |
15.93 |
|
|
|
| 21 |
B2 |
1219 |
1153 |
11.36 |
|
|
|
| 22 |
B2 |
1064 |
1006 |
318.38 |
|
|
|
| 23 |
B2 |
756 |
715 |
1.54 |
|
|
|
| 24 |
B2 |
503 |
476 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16616.4 cm
-1
Scaled (by 0.9454) Zero Point Vibrational Energy (zpe) 15709.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.870 |
| O2 |
0.000 |
0.000 |
2.063 |
| O3 |
0.000 |
1.137 |
0.056 |
| O4 |
0.000 |
-1.137 |
0.056 |
| C5 |
0.000 |
0.774 |
-1.294 |
| C6 |
0.000 |
-0.774 |
-1.294 |
| H7 |
-0.888 |
1.187 |
-1.774 |
| H8 |
0.888 |
1.187 |
-1.774 |
| H9 |
0.888 |
-1.187 |
-1.774 |
| H10 |
-0.888 |
-1.187 |
-1.774 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1926 | 1.3980 | 1.3980 | 2.2988 | 2.2988 | 3.0314 | 3.0314 | 3.0314 | 3.0314 |
O2 | 1.1926 | | 2.3062 | 2.3062 | 3.4452 | 3.4452 | 4.1132 | 4.1132 | 4.1132 | 4.1132 | O3 | 1.3980 | 2.3062 | | 2.2731 | 1.3980 | 2.3400 | 2.0347 | 2.0347 | 3.0882 | 3.0882 | O4 | 1.3980 | 2.3062 | 2.2731 | | 2.3400 | 1.3980 | 3.0882 | 3.0882 | 2.0347 | 2.0347 | C5 | 2.2988 | 3.4452 | 1.3980 | 2.3400 | | 1.5489 | 1.0902 | 1.0902 | 2.2059 | 2.2059 | C6 | 2.2988 | 3.4452 | 2.3400 | 1.3980 | 1.5489 | | 2.2059 | 2.2059 | 1.0902 | 1.0902 | H7 | 3.0314 | 4.1132 | 2.0347 | 3.0882 | 1.0902 | 2.2059 | | 1.7754 | 2.9647 | 2.3743 | H8 | 3.0314 | 4.1132 | 2.0347 | 3.0882 | 1.0902 | 2.2059 | 1.7754 | | 2.3743 | 2.9647 | H9 | 3.0314 | 4.1132 | 3.0882 | 2.0347 | 2.2059 | 1.0902 | 2.9647 | 2.3743 | | 1.7754 | H10 | 3.0314 | 4.1132 | 3.0882 | 2.0347 | 2.2059 | 1.0902 | 2.3743 | 2.9647 | 1.7754 | |
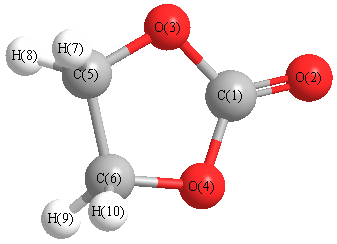 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.601 |
|
C1 |
O4 |
C6 |
110.601 |
| O2 |
C1 |
O3 |
125.613 |
|
O2 |
C1 |
O4 |
125.613 |
| O3 |
C1 |
O4 |
108.775 |
|
O3 |
C5 |
C6 |
105.012 |
| O3 |
C5 |
H7 |
109.087 |
|
O3 |
C5 |
H8 |
109.087 |
| O4 |
C6 |
C5 |
105.012 |
|
O4 |
C6 |
H9 |
109.087 |
| O4 |
C6 |
H10 |
109.087 |
|
C5 |
C6 |
H9 |
112.245 |
| C5 |
C6 |
H10 |
112.245 |
|
C6 |
C5 |
H7 |
112.245 |
| C6 |
C5 |
H8 |
112.245 |
|
H7 |
C5 |
H8 |
109.027 |
| H9 |
C6 |
H10 |
109.027 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -341.512008 |
| Energy at 298.15K | -341.518441 |
| HF Energy | -340.537699 |
| Nuclear repulsion energy | 245.230696 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3194 |
3019 |
16.48 |
|
|
|
| 2 |
A |
3111 |
2942 |
30.65 |
|
|
|
| 3 |
A |
1933 |
1828 |
622.06 |
|
|
|
| 4 |
A |
1585 |
1498 |
4.09 |
|
|
|
| 5 |
A |
1443 |
1364 |
9.74 |
|
|
|
| 6 |
A |
1282 |
1212 |
15.99 |
|
|
|
| 7 |
A |
1230 |
1162 |
148.59 |
|
|
|
| 8 |
A |
1179 |
1115 |
70.88 |
|
|
|
| 9 |
A |
980 |
927 |
7.03 |
|
|
|
| 10 |
A |
824 |
779 |
46.41 |
|
|
|
| 11 |
A |
721 |
681 |
0.66 |
|
|
|
| 12 |
A |
171 |
162 |
0.92 |
|
|
|
| 13 |
B |
3203 |
3028 |
23.63 |
|
|
|
| 14 |
B |
3115 |
2945 |
42.97 |
|
|
|
| 15 |
B |
1576 |
1490 |
3.45 |
|
|
|
| 16 |
B |
1433 |
1354 |
13.41 |
|
|
|
| 17 |
B |
1290 |
1219 |
6.96 |
|
|
|
| 18 |
B |
1193 |
1127 |
9.48 |
|
|
|
| 19 |
B |
1059 |
1001 |
284.47 |
|
|
|
| 20 |
B |
924 |
873 |
14.08 |
|
|
|
| 21 |
B |
767 |
725 |
27.70 |
|
|
|
| 22 |
B |
684 |
647 |
2.30 |
|
|
|
| 23 |
B |
503 |
476 |
0.07 |
|
|
|
| 24 |
B |
140 |
132 |
2.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16769.3 cm
-1
Scaled (by 0.9454) Zero Point Vibrational Energy (zpe) 15853.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31+G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.864 |
| O2 |
0.000 |
0.000 |
2.056 |
| O3 |
0.000 |
1.136 |
0.043 |
| O4 |
0.000 |
-1.136 |
0.043 |
| C5 |
0.223 |
0.733 |
-1.281 |
| C6 |
-0.223 |
-0.733 |
-1.281 |
| H7 |
-0.363 |
1.362 |
-1.947 |
| H8 |
1.285 |
0.834 |
-1.522 |
| H9 |
0.363 |
-1.362 |
-1.947 |
| H10 |
-1.285 |
-0.834 |
-1.522 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1924 | 1.4019 | 1.4019 | 2.2778 | 2.2778 | 3.1450 | 2.8348 | 3.1450 | 2.8348 |
O2 | 1.1924 | | 2.3121 | 2.3121 | 3.4243 | 3.4243 | 4.2446 | 3.8919 | 4.2446 | 3.8919 | O3 | 1.4019 | 2.3121 | | 2.2724 | 1.4019 | 2.3012 | 2.0355 | 2.0466 | 3.2148 | 2.8247 | O4 | 1.4019 | 2.3121 | 2.2724 | | 2.3012 | 1.4019 | 3.2148 | 2.8247 | 2.0355 | 2.0466 | C5 | 2.2778 | 3.4243 | 1.4019 | 2.3012 | | 1.5319 | 1.0880 | 1.0932 | 2.2029 | 2.1875 | C6 | 2.2778 | 3.4243 | 2.3012 | 1.4019 | 1.5319 | | 2.2029 | 2.1875 | 1.0880 | 1.0932 | H7 | 3.1450 | 4.2446 | 2.0355 | 3.2148 | 1.0880 | 2.2029 | | 1.7822 | 2.8199 | 2.4194 | H8 | 2.8348 | 3.8919 | 2.0466 | 2.8247 | 1.0932 | 2.1875 | 1.7822 | | 2.4194 | 3.0631 | H9 | 3.1450 | 4.2446 | 3.2148 | 2.0355 | 2.2029 | 1.0880 | 2.8199 | 2.4194 | | 1.7822 | H10 | 2.8348 | 3.8919 | 2.8247 | 2.0466 | 2.1875 | 1.0932 | 2.4194 | 3.0631 | 1.7822 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
108.662 |
|
C1 |
O4 |
C6 |
108.662 |
| O2 |
C1 |
O3 |
125.857 |
|
O2 |
C1 |
O4 |
125.857 |
| O3 |
C1 |
O4 |
108.286 |
|
O3 |
C5 |
C6 |
103.240 |
| O3 |
C5 |
H7 |
109.025 |
|
O3 |
C5 |
H8 |
109.598 |
| O4 |
C6 |
C5 |
103.240 |
|
O4 |
C6 |
H9 |
109.025 |
| O4 |
C6 |
H10 |
109.598 |
|
C5 |
C6 |
H9 |
113.376 |
| C5 |
C6 |
H10 |
111.789 |
|
C6 |
C5 |
H7 |
113.376 |
| C6 |
C5 |
H8 |
111.789 |
|
H7 |
C5 |
H8 |
109.582 |
| H9 |
C6 |
H10 |
109.582 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
