return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N2O2 (Oxalamide)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-337.738252
Energy at 298.15K-337.744124
HF Energy-336.756704
Nuclear repulsion energy232.460868
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3780 3627 0.00      
2 Ag 3626 3479 0.00      
3 Ag 1847 1772 0.00      
4 Ag 1617 1551 0.00      
5 Ag 1449 1390 0.00      
6 Ag 1112 1067 0.00      
7 Ag 787 755 0.00      
8 Ag 538 516 0.00      
9 Ag 406 389 0.00      
10 Au 679 652 3.48      
11 Au 459 440 52.76      
12 Au 232 222 387.83      
13 Au 39 38 18.21      
14 Bg 839 805 0.00      
15 Bg 657 630 0.00      
16 Bg 180 173 0.00      
17 Bu 3781 3627 167.26      
18 Bu 3627 3480 138.16      
19 Bu 1808 1735 618.80      
20 Bu 1599 1534 295.64      
21 Bu 1332 1277 133.39      
22 Bu 1100 1055 5.08      
23 Bu 577 554 25.32      
24 Bu 281 269 34.83      

Unscaled Zero Point Vibrational Energy (zpe) 16174.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15517.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.18672 0.12339 0.07429

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.775 0.000
C2 0.000 -0.775 0.000
O3 1.040 1.424 0.000
O4 -1.040 -1.424 0.000
N5 -1.258 1.272 0.000
N6 1.258 -1.272 0.000
H7 -1.405 2.272 0.000
H8 -2.033 0.619 0.000
H9 1.405 -2.272 0.000
H10 2.033 -0.619 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 O4 N5 N6 H7 H8 H9 H10
C11.54951.22562.43201.35232.40192.05352.03893.35542.4649
C21.54952.43201.22562.40191.35233.35542.46492.05352.0389
O31.22562.43203.52602.30242.70422.58773.17623.71422.2717
O42.43201.22563.52602.70422.30243.71422.27172.58773.1762
N51.35232.40192.30242.70423.57691.01151.01324.43273.7951
N62.40191.35232.70422.30243.57694.43273.79511.01151.0132
H72.05353.35542.58773.71421.01154.43271.76835.34324.4922
H82.03892.46493.17622.27171.01323.79511.76834.49224.2503
H93.35542.05353.71422.58774.43271.01155.34324.49221.7683
H102.46492.03892.27173.17623.79511.01324.49224.25031.7683

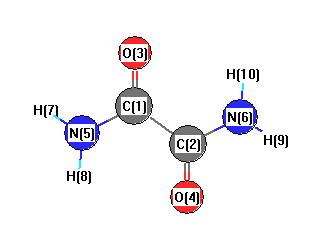
picture of Oxalamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 121.981 C1 C2 N6 111.551
C1 N5 H7 119.926 C1 N5 H8 118.367
C2 C1 O3 121.981 C2 C1 N5 111.551
C2 N6 H9 119.926 C2 N6 H10 118.367
O3 C1 N5 126.467 O4 C2 N6 126.467
H7 N5 H8 121.707 H9 N6 H10 121.707
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability