return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Furan, 2,5-dihydro-)

using model chemistry: QCISD/6-31G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G
 hartrees
Energy at 0K-230.203458
Energy at 298.15K-230.210567
HF Energy-229.681633
Nuclear repulsion energy170.181827
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3213 3096 11.87      
2 A1 3014 2904 9.72      
3 A1 1675 1614 0.35      
4 A1 1579 1521 0.16      
5 A1 1386 1336 0.52      
6 A1 1155 1113 1.01      
7 A1 987 951 6.51      
8 A1 832 801 8.62      
9 A1 736 710 8.77      
10 A2 3043 2932 0.00      
11 A2 1196 1152 0.00      
12 A2 1092 1052 0.00      
13 A2 919 885 0.00      
14 A2 383 369 0.00      
15 B1 3047 2936 96.06      
16 B1 1154 1112 6.59      
17 B1 1053 1015 16.00      
18 B1 685 660 37.05      
19 B1 116 112 14.20      
20 B2 3184 3068 3.32      
21 B2 3009 2900 98.76      
22 B2 1571 1514 2.35      
23 B2 1382 1332 2.80      
24 B2 1349 1300 1.80      
25 B2 1045 1007 68.31      
26 B2 925 892 3.99      
27 B2 798 769 4.44      

Unscaled Zero Point Vibrational Energy (zpe) 20263.4 cm-1
Scaled (by 0.9636) Zero Point Vibrational Energy (zpe) 19525.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G
ABC
0.25390 0.25326 0.13329

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 1.225
C2 0.000 1.217 0.366
C3 0.000 0.676 -1.060
C4 0.000 -0.676 -1.060
C5 0.000 -1.217 0.366
H6 0.897 1.815 0.601
H7 -0.897 1.815 0.601
H8 -0.897 -1.815 0.601
H9 0.897 -1.815 0.601
H10 0.000 1.323 -1.937
H11 0.000 -1.323 -1.937

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.49002.38242.38241.49002.11852.11852.11852.11853.42783.4278
C21.49001.52492.36942.43451.10331.10333.17083.17082.30523.4288
C32.38241.52491.35102.36942.20472.20473.12483.12481.09062.1828
C42.38242.36941.35101.52493.12483.12482.20472.20472.18281.0906
C51.49002.43452.36941.52493.17083.17081.10331.10333.42882.3052
H62.11851.10332.20473.12483.17081.79434.04903.62972.73654.1345
H72.11851.10332.20473.12483.17081.79433.62974.04902.73654.1345
H82.11853.17083.12482.20471.10334.04903.62971.79434.13452.7365
H92.11853.17083.12482.20471.10333.62974.04901.79434.13452.7365
H103.42782.30521.09062.18283.42882.73652.73654.13454.13452.6465
H113.42783.42882.18281.09062.30524.13454.13452.73652.73652.6465

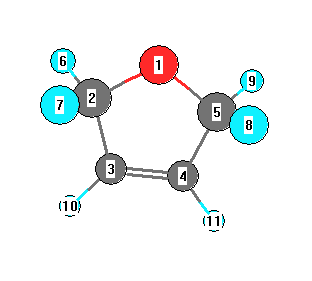
picture of Furan, 2,5-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 104.410 O1 C2 H6 108.633
O1 C2 H7 108.633 O1 C5 C4 104.410
O1 C5 H8 108.633 O1 C5 H9 108.633
C2 O1 C5 109.562 C2 C3 C4 110.809
C2 C3 H10 122.756 C3 C2 H6 113.064
C3 C2 H7 113.064 C3 C4 C5 110.809
C3 C4 H11 126.435 C4 C3 H10 126.435
C4 C5 H8 113.064 C4 C5 H9 113.064
C5 C4 H11 122.756 H6 C2 H7 108.802
H8 C5 H9 108.802
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability