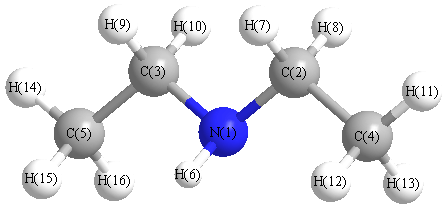
Vibrational Frequencies calculated at LSDA/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3389 |
3326 |
1.22 |
|
|
|
| 2 |
A' |
3092 |
3034 |
21.65 |
|
|
|
| 3 |
A' |
3067 |
3009 |
40.55 |
|
|
|
| 4 |
A' |
2989 |
2933 |
31.64 |
|
|
|
| 5 |
A' |
2973 |
2917 |
37.83 |
|
|
|
| 6 |
A' |
2834 |
2781 |
177.07 |
|
|
|
| 7 |
A' |
1492 |
1465 |
2.77 |
|
|
|
| 8 |
A' |
1463 |
1436 |
2.39 |
|
|
|
| 9 |
A' |
1449 |
1422 |
21.25 |
|
|
|
| 10 |
A' |
1395 |
1369 |
4.18 |
|
|
|
| 11 |
A' |
1348 |
1323 |
2.24 |
|
|
|
| 12 |
A' |
1279 |
1255 |
3.24 |
|
|
|
| 13 |
A' |
1210 |
1187 |
1.68 |
|
|
|
| 14 |
A' |
1148 |
1126 |
16.65 |
|
|
|
| 15 |
A' |
1083 |
1063 |
9.90 |
|
|
|
| 16 |
A' |
890 |
873 |
8.83 |
|
|
|
| 17 |
A' |
805 |
790 |
3.45 |
|
|
|
| 18 |
A' |
757 |
743 |
85.47 |
|
|
|
| 19 |
A' |
427 |
419 |
0.36 |
|
|
|
| 20 |
A' |
262 |
257 |
0.83 |
|
|
|
| 21 |
A' |
179 |
176 |
0.93 |
|
|
|
| 22 |
A' |
112 |
110 |
0.81 |
|
|
|
| 23 |
A" |
3092 |
3034 |
9.39 |
|
|
|
| 24 |
A" |
3067 |
3009 |
7.36 |
|
|
|
| 25 |
A" |
2989 |
2933 |
17.51 |
|
|
|
| 26 |
A" |
2974 |
2918 |
0.95 |
|
|
|
| 27 |
A" |
2830 |
2777 |
20.24 |
|
|
|
| 28 |
A" |
1479 |
1452 |
6.07 |
|
|
|
| 29 |
A" |
1464 |
1437 |
6.46 |
|
|
|
| 30 |
A" |
1457 |
1429 |
19.48 |
|
|
|
| 31 |
A" |
1434 |
1407 |
26.37 |
|
|
|
| 32 |
A" |
1370 |
1344 |
26.79 |
|
|
|
| 33 |
A" |
1311 |
1287 |
24.49 |
|
|
|
| 34 |
A" |
1241 |
1218 |
5.12 |
|
|
|
| 35 |
A" |
1205 |
1182 |
38.43 |
|
|
|
| 36 |
A" |
1091 |
1070 |
2.91 |
|
|
|
| 37 |
A" |
1056 |
1036 |
14.30 |
|
|
|
| 38 |
A" |
946 |
928 |
0.07 |
|
|
|
| 39 |
A" |
790 |
776 |
0.32 |
|
|
|
| 40 |
A" |
417 |
409 |
0.79 |
|
|
|
| 41 |
A" |
264 |
259 |
0.81 |
|
|
|
| 42 |
A" |
127 |
125 |
1.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32121.5 cm
-1
Scaled (by 0.9813) Zero Point Vibrational Energy (zpe) 31520.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.495 |
|
|
|
| 2 |
C |
-0.186 |
|
|
|
| 3 |
C |
-0.186 |
|
|
|
| 4 |
C |
-0.546 |
|
|
|
| 5 |
C |
-0.546 |
|
|
|
| 6 |
H |
0.300 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
| 9 |
H |
0.131 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.189 |
|
|
|
| 14 |
H |
0.175 |
|
|
|
| 15 |
H |
0.167 |
|
|
|
| 16 |
H |
0.189 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.769 |
0.221 |
0.000 |
0.800 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.560 |
1.709 |
0.000 |
| y |
1.709 |
-34.178 |
0.000 |
| z |
0.000 |
0.000 |
-32.856 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.043 |
1.709 |
0.000 |
| y |
1.709 |
-0.969 |
0.000 |
| z |
0.000 |
0.000 |
1.013 |
|
| Polar |
|---|
| 3z2-r2 | 2.025 |
|---|
| x2-y2 | 0.617 |
|---|
| xy | 1.709 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.041 |
0.102 |
0.000 |
| y |
0.102 |
7.466 |
0.000 |
| z |
0.000 |
0.000 |
9.489 |
<r2> (average value of r
2) Å
2
| <r2> |
184.074 |
| (<r2>)1/2 |
13.567 |